<?xml version="1.0" encoding="utf-8"?><feed xmlns="http://www.w3.org/2005/Atom" ><generator uri="https://jekyllrb.com/" version="3.10.0">Jekyll</generator><link href="https://animal-adna-research-community.github.io/AaRC.github.io//feed.xml" rel="self" type="application/atom+xml" /><link href="https://animal-adna-research-community.github.io/AaRC.github.io//" rel="alternate" type="text/html" /><updated>2026-09-24T11:15:44+00:00</updated><id>https://animal-adna-research-community.github.io/AaRC.github.io//feed.xml</id><title type="html">AaRC</title><subtitle>Animal
aDNA
Research
Community
</subtitle><author><name>AaRC</name></author><entry><title type="html">Interview to our July AaRCTikTalks speaker Molly Cassatt-Johnstone</title><link href="https://animal-adna-research-community.github.io/AaRC.github.io//blog/2026/09/12/Interview-July-Speaker1/" rel="alternate" type="text/html" title="Interview to our July AaRCTikTalks speaker Molly Cassatt-Johnstone" /><published>2026-09-12T00:00:00+00:00</published><updated>2026-09-12T00:00:00+00:00</updated><id>https://animal-adna-research-community.github.io/AaRC.github.io//blog/2026/09/12/Interview-July-Speaker1</id><content type="html" xml:base="https://animal-adna-research-community.github.io/AaRC.github.io//blog/2026/09/12/Interview-July-Speaker1/"><![CDATA[<h2 id="july-2026">July 2026</h2>
<h1 id="the-true-life-of-the-american-cheetah">The true life of the American cheetah</h1>
<p><em>“For animals belonging to two most distinct lines of descent, may readily become adapted to similar conditions, and thus assume a close external resemblance; but such resemblances will not reveal, will rather tend to conceal their blood-relationship to their proper lines of descent.”</em> (Charles Darwin, On the Origin of Species, 1859).</p>

<p> </p>

<h2 id="a-cat-named-after-a-look">A cat named after a look</h2>
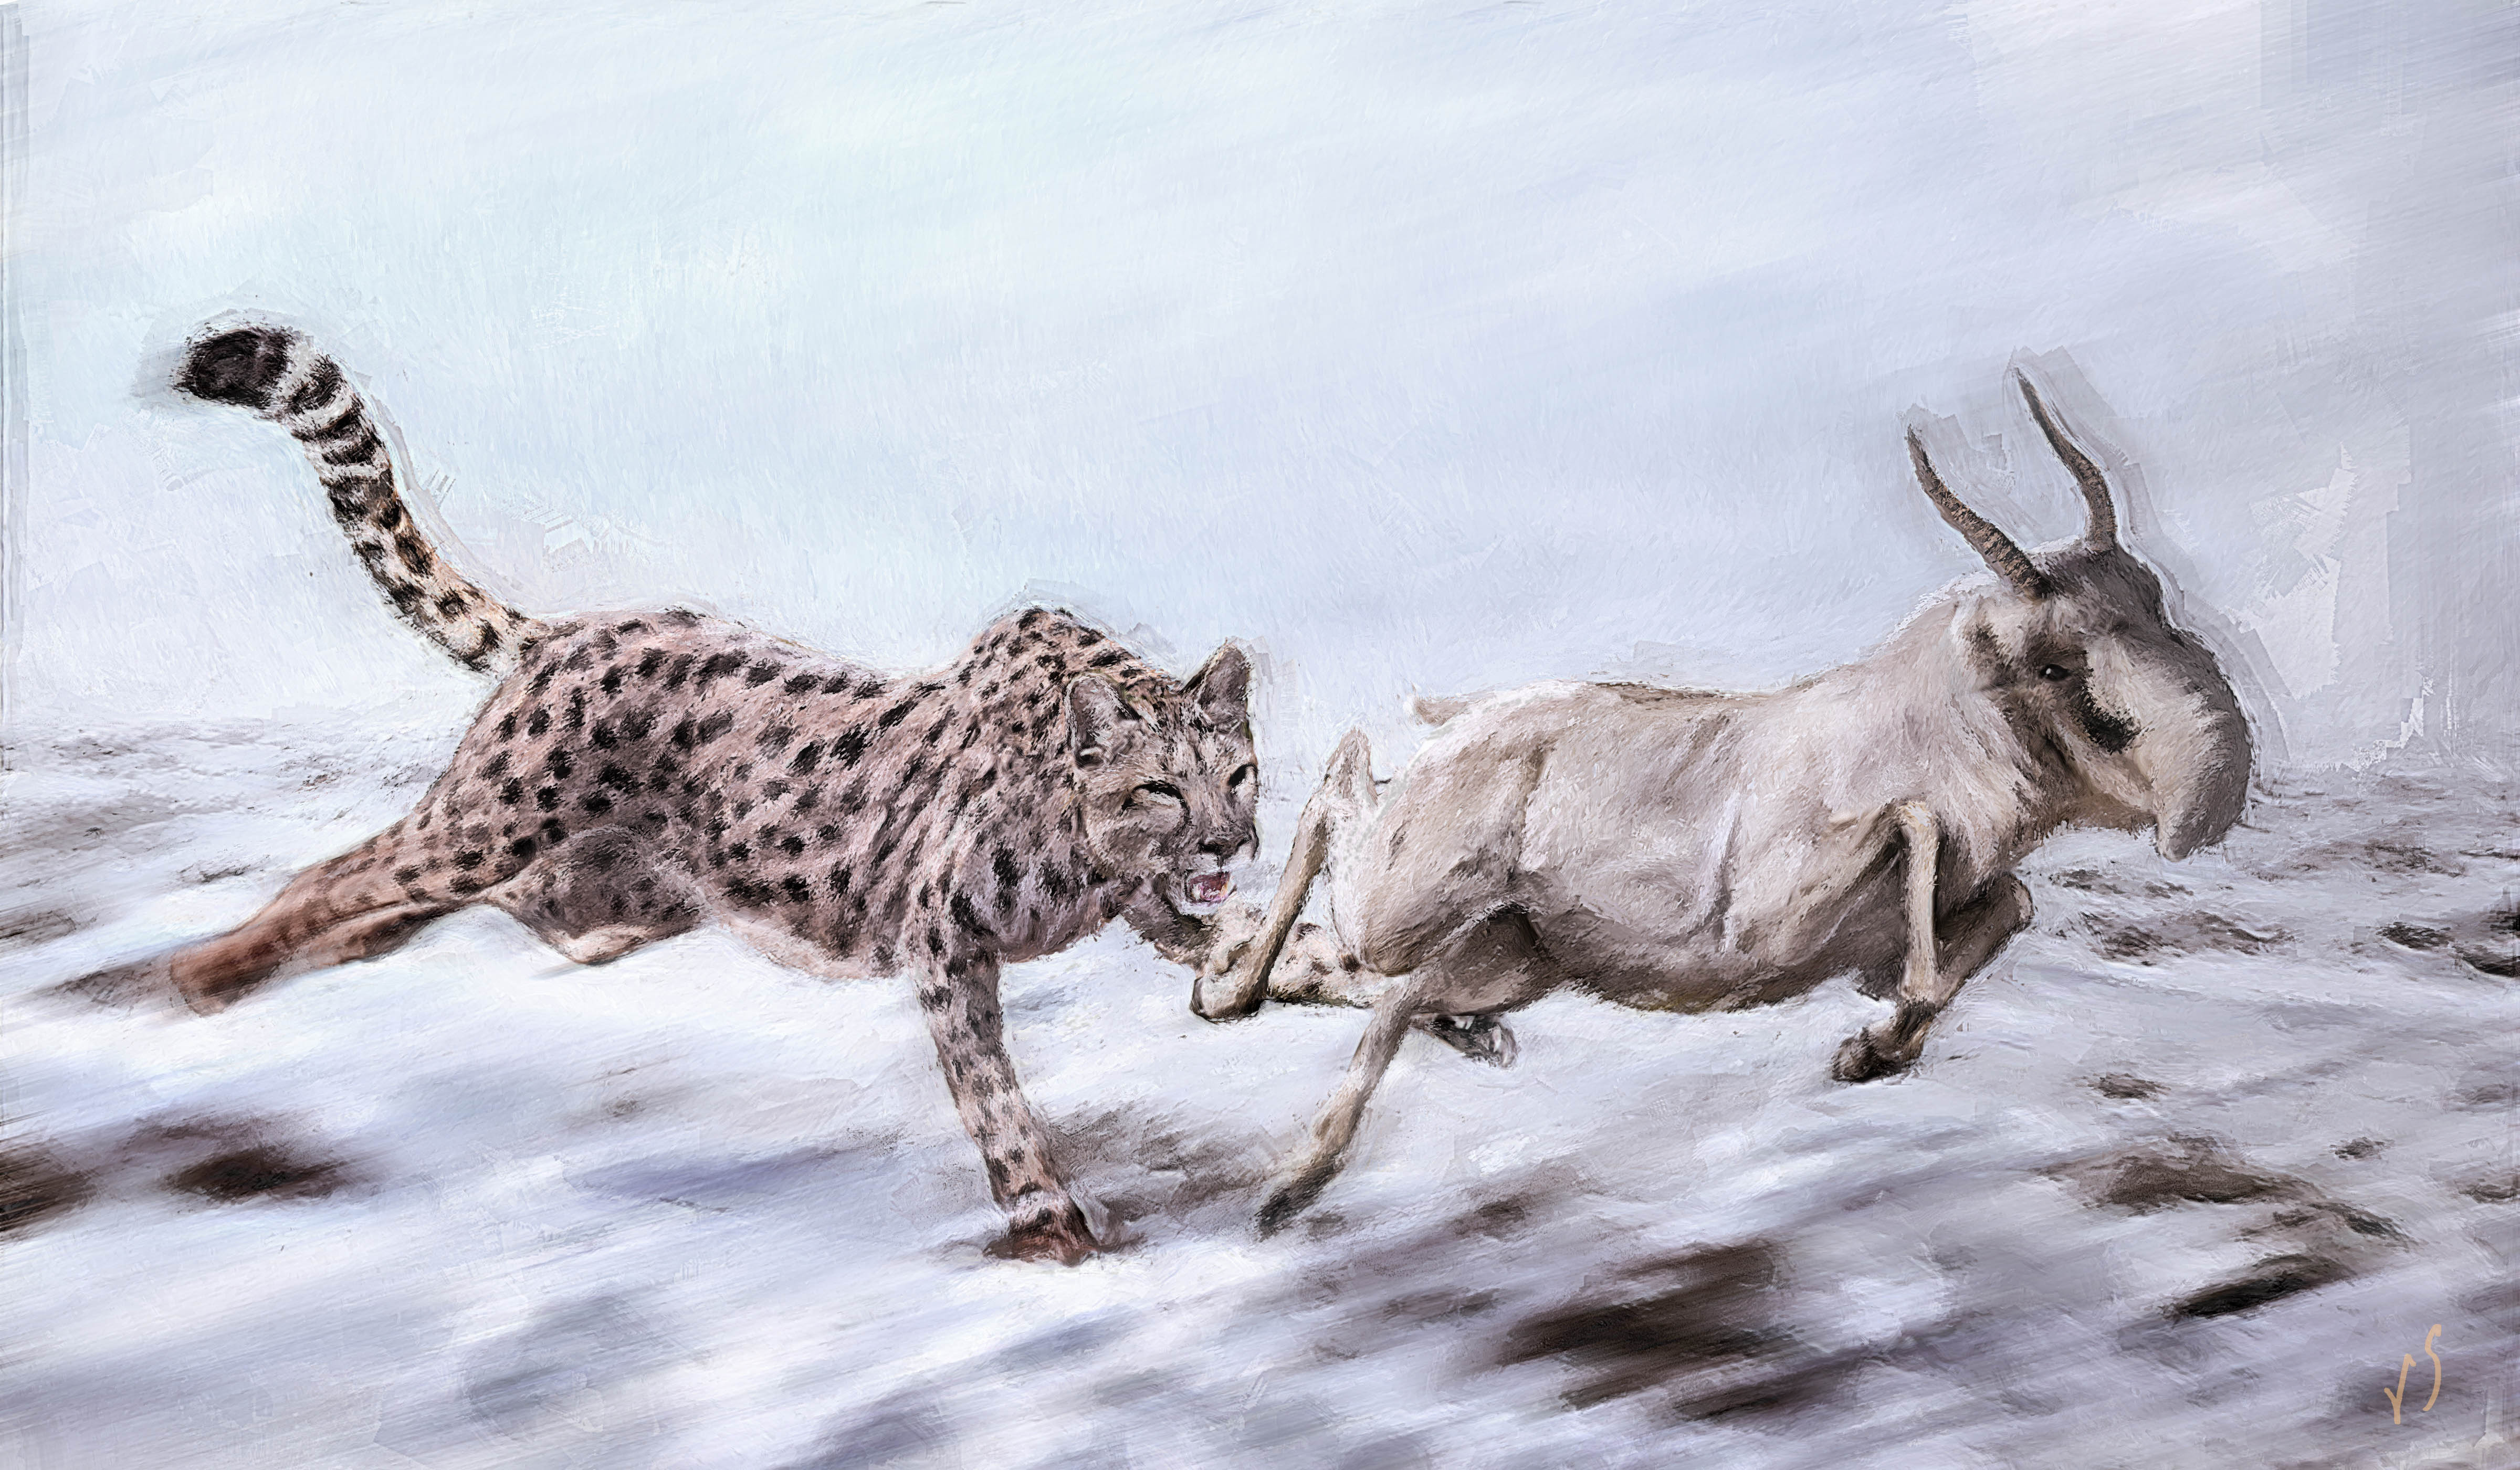
<p>Pleistocene North America held a far richer set of big cats than the continent has today: alongside pumas, jaguars, and lynxes, there were American lions, saber-toothed cats, and the lesser-known American cheetah, <em>Miracinonyx trumani</em>. As our AaRC TikTalk speaker Molly Cassatt-Johnstone explained, the animal was described in 1979 and named for its resemblance to the Old World cheetah, <em>Acinonyx</em> [<strong>1</strong>]. The two shared a sprinter’s build with long front legs, a short muzzle, and enlarged nasal passages for taking in oxygen during a chase. The similarity was such that researchers thought anything other than shared ancestry between both species would be hard to explain. Indeed, Miracinonyx was cast as a specialist pursuit predator of the American pronghorn, an image so established that paleo-art still shows it mid-chase. As it turned out, they were mistaken.</p>

<p> </p>

<p><img src="/assets/media/Blog_July_2026_MCJ_Figure.jpg" alt="Cheetah chasing" title="Cheetah chasing" /></p>

<p><em>A depiction of Miracinonyx trumani racing after its pray. Illustration by Velizar Simeonovski.</em></p>
<p> </p>

<h2 id="cracks-in-the-cheetah-story">Cracks in the cheetah story</h2>
<p>The first cracks were anatomical. As Molly described, fossils found in the Grand Canyon in the 1930s were later reassigned to <em>Miracinonyx</em>, which was surprising, since there is little open ground to sprint across in a canyon. An animal living there would have been climbing dry uplands and cliffs after bighorn, closer to the ambush hunting of a snow leopard than the pursuit of a cheetah. Later morphology work pointed the same way.  Studies on the elbow-joint shape, a useful indicator of hunting style, placed Miracinonyx between the puma and the cheetah [<strong>2</strong>], while a study of wrist bones grouped it with the large pantherine cats [<strong>3</strong>]. As our TikTalk speaker Molly argues, these cats probably kept an intermediate lifestyle, able to grapple and climb while still carrying some sprinter’s anatomy, occupying a niche with no clear modern equivalent.
 </p>

<h2 id="what-the-genome-revealed">What the genome revealed</h2>
<p>Early hypotheses placed <em>Miracinonyx</em> as a close relative to the African cheetah, but in 2005 Ross Barnett et al. analyzed mitochondrial DNA fragments that instead made it closest to the puma [<strong>4</strong>]. Because cats are known for mismatches between their mitochondrial and nuclear genomes, in her recently published work Molly aimed to test such evolutionary origin with nuclear genomes [<strong>5</strong>]. Her own project began with three fossils from Yukon, Canada, labelled as pumas and dated to around 30,000 years ago. The first phylogeny looked nothing like a puma, and her initial assumption was that she had made an error. A fourth sample though, confirmed as <em>Miracinonyx</em> from Natural Trap Cave in Wyoming and dated to about 23,000 years ago, clustered with all three Yukon “pumas”. Sequencing the two best samples to high coverage confirmed that <em>Miracinonyx</em> was indeed a sister species to the puma, with both lineages splitting around 2.6 million years ago [<strong>5</strong>]. This is also the most evident explanation, since a cheetah relationship would have required an awkward back-dispersal across continents.
 </p>

<h2 id="a-cat-at-two-ends-of-a-continent">A cat at two ends of a continent</h2>
<p>The Yukon fossils extended <em>Miracinonyx</em>’s known range far to the north. The earlier record tied this species to the grasslands of the temperate North America, where pronghorn lived [<strong>6</strong>], but Molly’s Beringian samples came from far northern regions. Furthermore, stable isotopes extracted from these bones were able to explain what they ate, and the answer was unexpected. The Wyoming samples resembled other Pleistocene carnivores, but the three Yukon samples sat far away from any other profile of Ice Age carnivore measured. Compared against modern salmon-eating brown bears, marine-feeding “sea wolves” of coastal British Columbia, and fish-eating orcas, the Yukon <em>Miracinonyx</em> looked like having a diet made up almost entirely of aquatic resources [<strong>5</strong>]. The same species appeared to be both a grassland hunter at one end of its range and a fish specialist at the other. Besides, their genomes showed low diversity, close to African cheetahs and inbred Florida panthers, but spread evenly across the genome, a signature revealing a small population held steadily over a long time rather than a sudden population bottleneck. Indeed, effective population size modeling confirmed a slow, sustained decline into the late Pleistocene [<strong>5</strong>].
 </p>

<h2 id="why-reading-species-on-their-own-terms-matters">Why reading species on their own terms matters</h2>
<p>The thread running through Molly’s talk is a caution about analogy. We picture extinct animals by comparing them to living ones, and the risk is that the comparison becomes the conclusion, reducing the animal to a copy of whatever it resembles nowadays. For decades a cheetah-shaped cat was assumed to have lived a cheetah’s life. What emerged instead was a puma relative with wide ecological range up to the Arctic, where aquatic hunting became the way to survive. As Molly put it, interpreting the past through the lens of the present can strip an animal of its real dimensionality, which is precisely why her data is so valuable. A shared body shape is not a shared way of life.</p>

<p> </p>

<h2 id="references">References</h2>
<ol>
  <li><a href="https://www.science.org/doi/10.1126/science.205.4411.1155">Adams, D. B. The cheetah: Native American. Science (1979). 205, 1155–1158 (1979).</a></li>
  <li><a href="https://royalsocietypublishing.org/rsbl/article/19/1/20220483/88047/Elbow-joint-morphology-in-the-North-American">Figueirido, B. et al. Elbow-joint morphology in the North American ‘cheetah-like’ cat <em>Miracinonyx trumani</em>. Biol Lett 19, (2023).</a></li>
  <li><a href="https://onlinelibrary.wiley.com/doi/abs/10.1002/jmor.21024">Dunn, R. H., Cooper, C., Lemert, J., Mironov, N. &amp; Meachen, J. A. Locomotor correlates of the scapholunar of living and extinct carnivorans. J Morphol 280, 1197–1206 (2019).</a></li>
  <li><a href="https://www.cell.com/current-biology/fulltext/S0960-9822(05)00836-5">Barnett, R. et al. Evolution of the extinct Sabretooths and the American cheetah-like cat. Curr Biol 15, (2005).</a></li>
  <li><a href="https://www.sciencedirect.com/science/article/pii/S0960982226010031">Cassatt-Johnstone, M. et al. Range and diet diversity in the Pleistocene American “cheetah,” <em>Miracinonyx trumani</em>. Curr Biol S0960-9822(26)01003-1 (2026).</a></li>
  <li><a href="https://www.tandfonline.com/doi/abs/10.1080/02724634.1990.10011827">Van Valkenburgh, B., Grady, F. &amp; Kurtén, B. The Plio-Pleistocene cheetah-like cat Miracinonyx inexpectatus of North America. J Vertebr Paleontol 10, 434–454 (1990).</a></li>
</ol>

<p> </p>

<p>Below, Molly shared with us further details about her profile, career, prospects and future projects:
 </p>

<p><strong>1. Briefly introduce yourself. What is your origin story for how you got into science?</strong> <br />
I did my undergraduate degree in Cognitive Science, thinking I would perhaps pursue clinical research or work in science communication and education. However, my final quarter of my undergraduate degree I took a field course with Dr. Beth Shapiro that reshaped my career intentions. The course led to a summer internship in her lab, which conveniently helped me sort out what it meant to be a college graduate and start a career. I had no prior experience with molecular biology, aside from general intro biology classes, so the internship was a very steep learning curve. I realized I loved the challenge of learning all this new science in a very applied manner. I was subsequently offered a full-time technician position where I could continue to build the foundational knowledge I had been missing in molecular biology while exploring how this information can be used to answer evolutionary questions. I’d always thought I liked ecology, but during that internship I realized the questions I was interested in were actually all evolutionary.
 </p>

<p><strong>2. How and/or why did you start working on this project?</strong> <br />
This project was actually started with three fossils from the Yukon, Canada that had been sent to Dr. Shapiro’s lab as part of a different student’s dissertation project. The fossils were originally identified as pumas, all dated to around 30,000 years ago. These samples would have been paramount for understanding the evolutionary history of pumas in North America, as they predate the coalescent time of all North American puma lineages. However, when we first screened the samples and realized they sat far outside any known puma mitochondrial diversity, the PhD student whose project it had been didn’t have the time to take this on, so I was lucky enough to get handed this as a project and got to see it through from the first surprising result to the end, and the results have finally been published!
 </p>

<p><strong>3. Were there any major challenges in this project? How did you overcome them?</strong> <br />
Definitely. For the first several years, we only had the Yukon samples, and while confirming they represented something far outside known puma diversity was exciting, I struggled with how to build that into a compelling scientific story. It was easy enough to generate nuclear genomic data and document a range extension into Beringia, but for a while, the narrative gap was the biggest obstacle. I had an interesting discovery without a clear sense of the larger story it was telling. Thankfully, that changed when we gained access to the Natural Trap Cave samples. Having a second population to compare against transformed the project, where instead of describing one isolated, unusual specimen, I could start asking comparative questions, which really helped to frame the manuscript into what I think is a much more compelling paper.
 </p>

<p><strong>4. What do you think are the main take-home messages of this project?</strong> <br />
The take-home message for me was that Miracinonyx trumani wasn’t behaving as a single predator across its range; instead, we see evidence in Beringia that they were using a strikingly different ecological strategy than we’d expect from a cursorial, cheetah-like hunter. We see this kind of ecological flexibility constantly in modern carnivores but it’s easy, at least for me, to forget that extinct species were capable of the same behavioral and ecological plasticity. There’s a natural tendency to flatten extinct animals into a single fixed caricature, in part because the fossil record so often gives us just one snapshot to work with. This project was a reminder that we owe extinct species the same dynamism and variability we readily accept in extant species. That’s also why multi-proxy approaches and strong collaborations are so important. A single data type will always struggle to capture the total complexity of an organism, living or extinct.
 </p>

<p><strong>5. What do you think is missing in the field that you would like to work on?</strong> <br />
What a big question! I think we’re seeing a lot more temporal genomics recently, using historical or museum specimens to add in a pre-Anthropocene baseline to conservation genomics studies. I also think temporal sampling has real, underused potential to reduce the burden we place on wild and threatened populations. I’m excited to work within that sector, answering conservation questions while minimizing invasive approaches. As wet lab and sequencing costs continue to drop, figuring out how to further reduce the cost, both in resources and in harm to wild populations, is something I think is very valuable to the field.
 </p>

<p><strong>6. Where do you see yourself in the near future?</strong> <br />
I’m working toward finishing my PhD, so the future I’m envisioning is a little uncertain! I am trying to envision having a very stress-free and completely relaxed defense, and then starting the next adventure, whatever that may be! I definitely see a lot of applications in my future, so I’m very open to anyone who’s interested in collaborating or furthering opportunities!
 </p>

<p><strong>7. Free space to tell something you would like to remark.</strong> <br />
Thanks so much for having me! Excited to be a part of this community!</p>]]></content><author><name>AaRC</name></author><category term="Blog" /><summary type="html"><![CDATA[July 2026 The true life of the American cheetah “For animals belonging to two most distinct lines of descent, may readily become adapted to similar conditions, and thus assume a close external resemblance; but such resemblances will not reveal, will rather tend to conceal their blood-relationship to their proper lines of descent.” (Charles Darwin, On the Origin of Species, 1859).]]></summary></entry><entry><title type="html">Interview to our July AaRCTikTalks speaker Michael Campana</title><link href="https://animal-adna-research-community.github.io/AaRC.github.io//blog/2026/09/12/Interview-July-Speaker2/" rel="alternate" type="text/html" title="Interview to our July AaRCTikTalks speaker Michael Campana" /><published>2026-09-12T00:00:00+00:00</published><updated>2026-09-12T00:00:00+00:00</updated><id>https://animal-adna-research-community.github.io/AaRC.github.io//blog/2026/09/12/Interview-July-Speaker2</id><content type="html" xml:base="https://animal-adna-research-community.github.io/AaRC.github.io//blog/2026/09/12/Interview-July-Speaker2/"><![CDATA[<h2 id="july-2026">July 2026</h2>
<h1 id="tracing-lions-decline-through-the-bones-of-roosevelt-expeditions">Tracing lion’s decline through the bones of Roosevelt expeditions</h1>
<p><em>“But the lions now offer, and have always offered, the chief source of unpleasant excitement.”</em> by Theodore Roosevelt (African Game Trails, 1910).</p>

<p> </p>

<h2 id="a-stronghold-under-pressure">A stronghold under pressure</h2>
<p>The lion was once among the most widespread large carnivores on Earth, ranging from the Cape of Good Hope to the Mediterranean and from West Africa to India. Today the species survives only in western India and Sub-Saharan Africa, and African populations have fallen dramatically over the last several decades, with an estimate of 20,000 to 25,000 individuals left [<strong>1</strong>]. Nevertheless, East Africa is still described as a modern lion stronghold, home of many of the continent’s largest populations [<strong>2</strong>]. A central question in conservation efforts of the lion species is how much of the genetic diversity and connectivity we see today reflects a truly healthy situation, and how much is already an eroded version of something richer that existed decades ago. To answer that though, we require a window into the past, something that museum collections can provide.</p>

<p> </p>

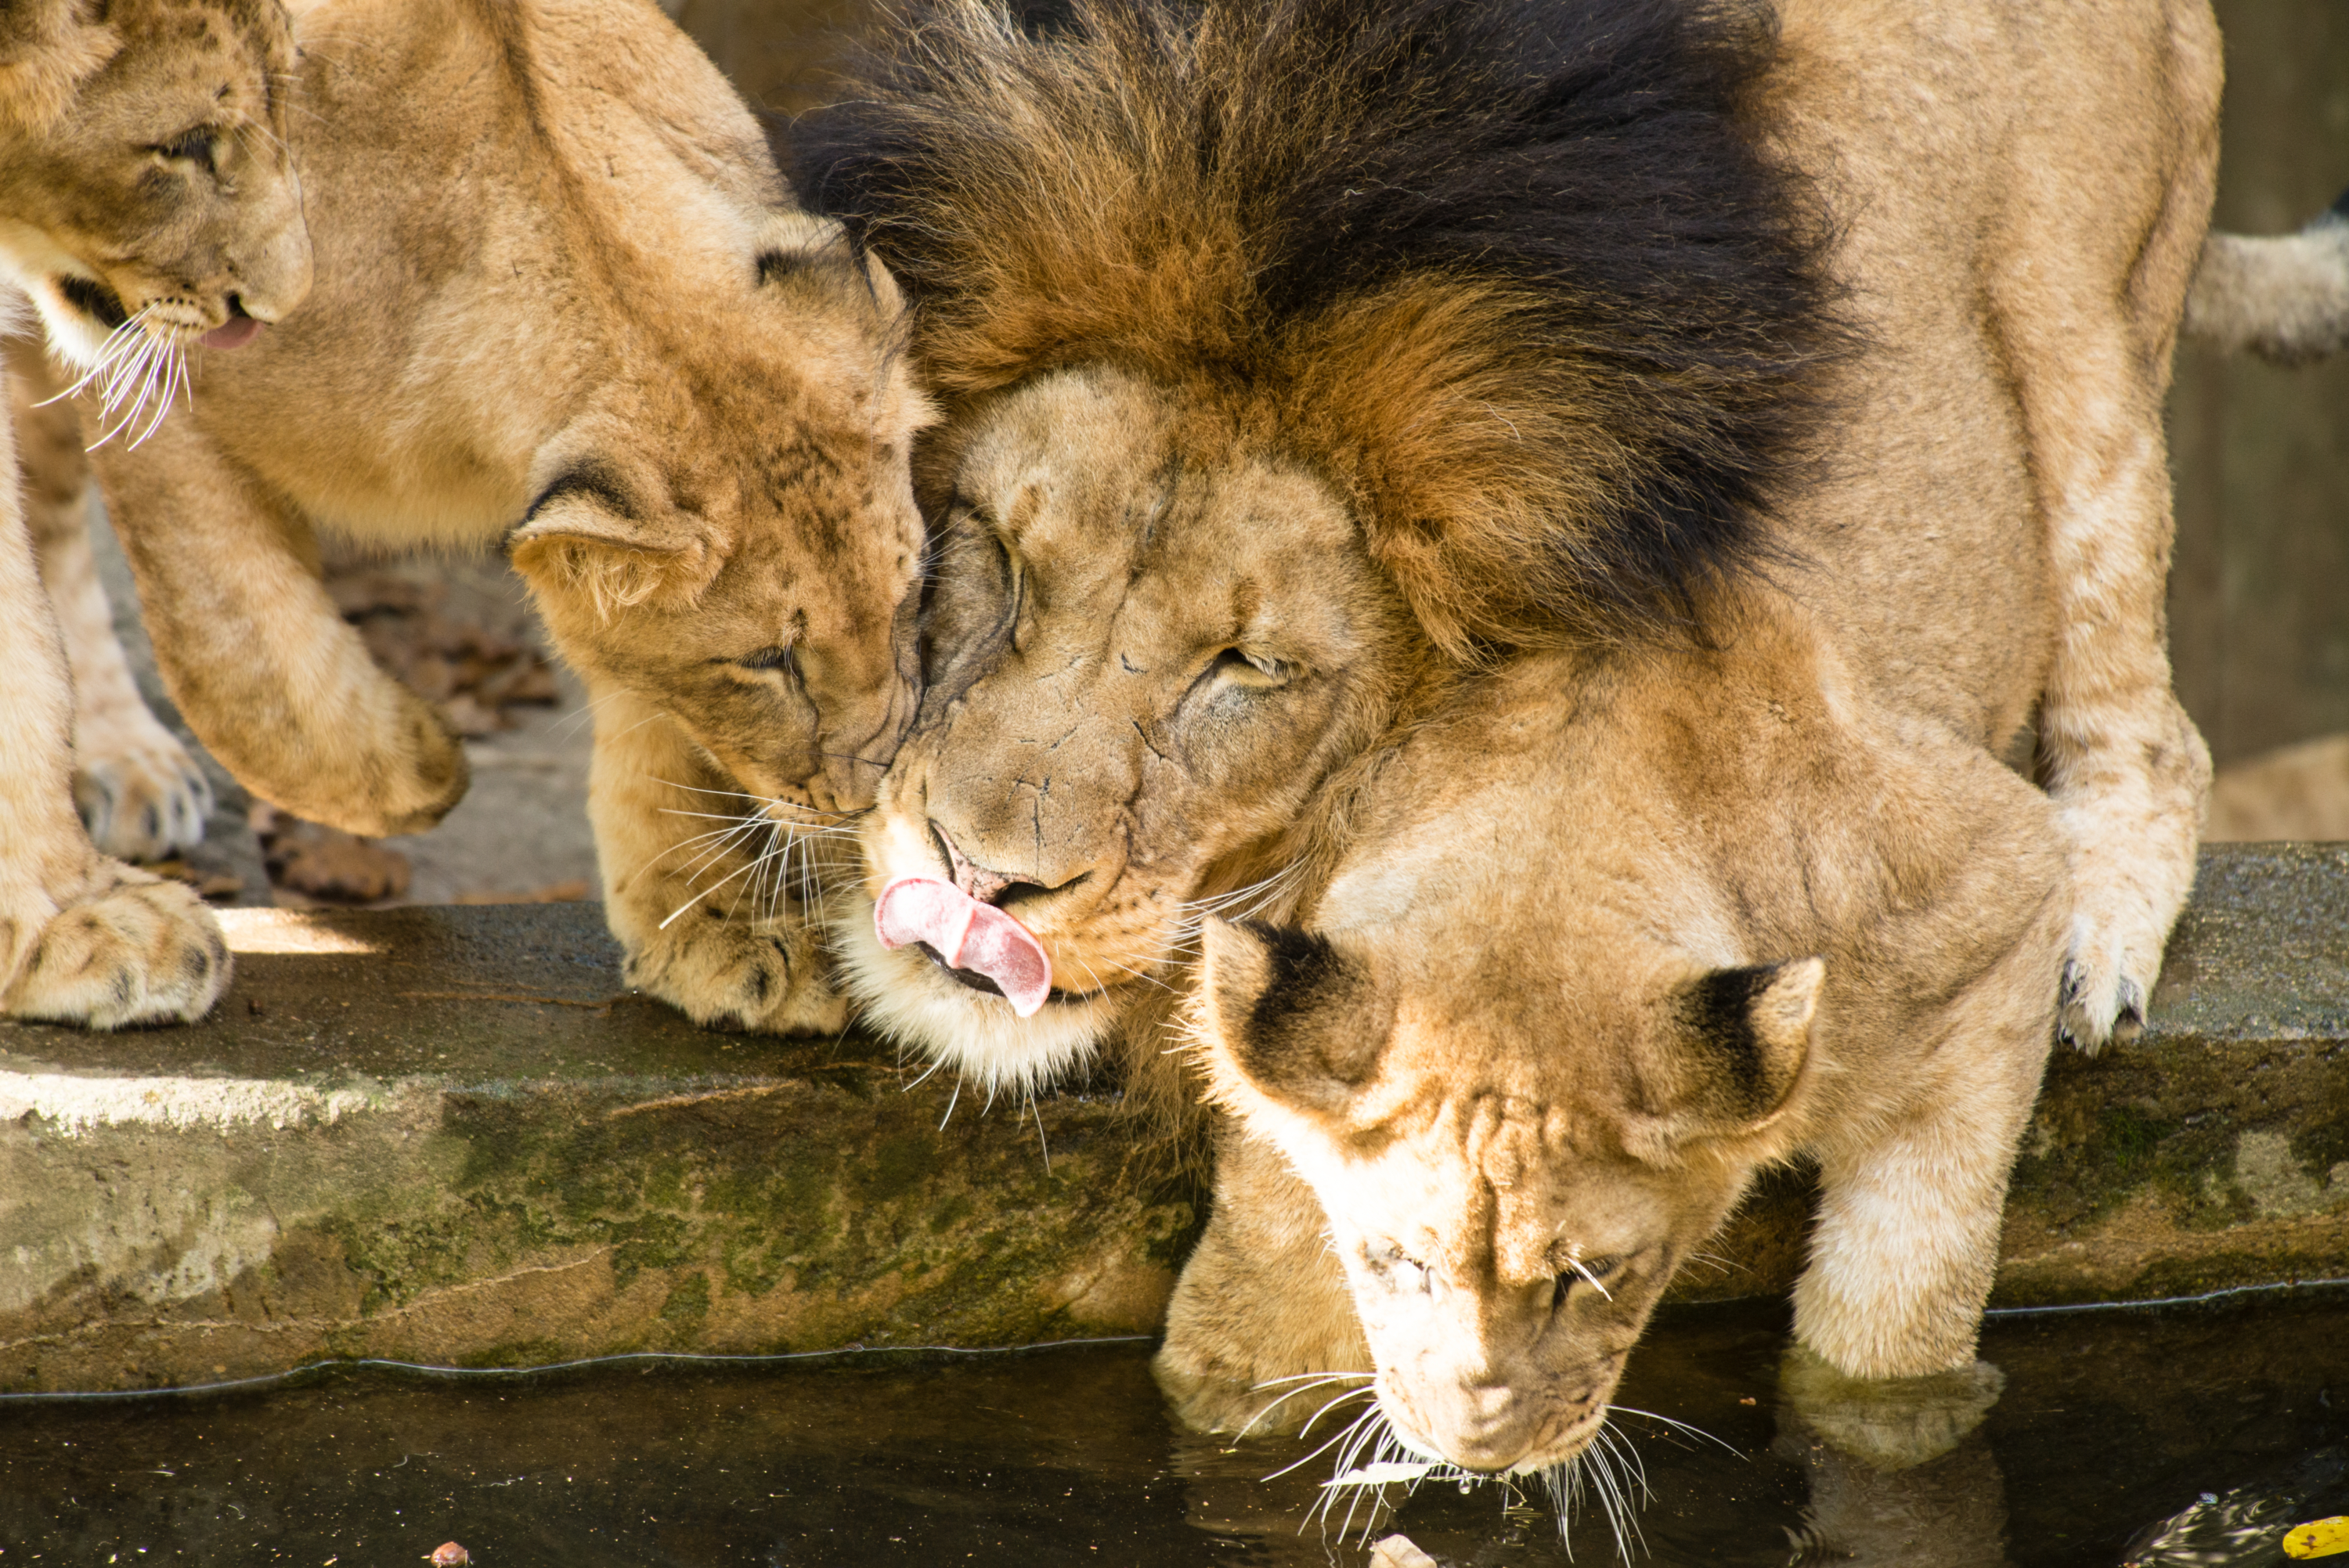
<p><img src="/assets/media/Blog_July_2026_MC_Figure.jpg" alt="Lions drinking" title="Lions drinking" /></p>

<p><em>African lions drinking in a pond. Photo by Connor Mallon, Smithsonian's National Zoo.</em></p>
<p> </p>

<h2 id="bones-and-skins-that-carry-a-genome">Bones and skins that carry a genome</h2>
<p>Our AaRC speaker, Dr. Michael Campana and his team made use of one such window in a direct way [<strong>3</strong>]. During the Smithsonian-Roosevelt East African Expedition of 1909 to 1911 and the Rainey Expedition of 1911 to 1912, thousands of lion specimens were collected from what is now Kenya and neighboring regions, and many of these have sat in the collections of the National Museum of Natural History ever since. The team sampled still attached soft tissue and previously damaged bone from these specimens.
In doing so, the authors managed to generate whole-genome data from 45 historical Kenyan lions from the Roosevelt and Rainey collections, added newly sequenced genomes from 94 recent Kenyan and Tanzanian lions, and combined everything with all publicly available East African sequence data. The final dataset came to 100 recent genomes, mostly from 1985 to 1999, and 102 historical genomes spanning 1896 to 1946. However, working with material this degraded is never clean or overly efficient, resulting in most historical genomes being sequenced at very low depth. To circumvent this, the team led by Dr. Campana leaned on genotype-likelihood methods rather than hard genotype calls.</p>

<p> </p>

<h2 id="what-the-genomes-revealed">What the genomes revealed</h2>
<p>Two signals came through clearly. First, principal component analysis and admixture modelling consistently separated recent Tanzanian lions, mostly from the Serengeti, from recent Kenyan lions, mostly from Tsavo East. When the historical individuals were added, they filled in a north-south cline between these two modern clusters, with historical Kenyan and Tanzanian lions sitting closer together than their present-day counterparts do. In other words, populations that a century ago were probably connected and interbreeding one another, now sit as more distinct, fragmented and isolated groups. A separate group was formed by three lions from the Aberdare mountain range, consistent with earlier evidence that this upland population is genetically distinctive [<strong>4</strong>].
The second signal is diversity loss. Recent East African lions were significantly more inbred than their historical counterparts, and the three deep-sequenced Roosevelt lions were significantly more heterozygous than recent Kenyan lions. Campana’s team also saw an accumulation of runs of homozygosity in recent genomes, although they treat that particular finding as suggestive rather than settled. The mitochondrial phylogeny, by contrast, showed widespread sharing of haplotypes across the continent and only shallow divergence, a reminder that maternally inherited markers can miss the fragmentation that whole genomes may identify [<strong>5</strong>].</p>

<p> </p>

<h2 id="a-historical-bottleneck-with-a-name">A historical bottleneck with a name</h2>
<p>Perhaps the most striking result is demographic. When the authors reconstructed effective population sizes for the Serengeti-Mara and Tsavo populations, they found a rapid decline beginning roughly 25 generations ago in both. At a lion generation time of about five years, that places the bottleneck in the late XIX century, coinciding with the great rinderpest panzootic that swept East Africa in the 1890s [<strong>6</strong>] and is estimated to have wiped out roughly 90% of the cattle and wild bovids in the region, with predators declining as their prey collapsed. Effective population size in Kenya was much larger, around 3,500, than in Tanzania, around 1,500, before the rinderpest bottleneck, while current effective sizes tell a sobering story, with only roughly 202 for the Serengeti-Mara and about 12 for Tsavo [<strong>3</strong>].</p>

<p> </p>

<h2 id="why-old-skins-still-matter">Why old skins still matter</h2>
<p>Reading genomes from lions shot more than a century ago, then laying them directly against lions alive today, is an experiment that no amount of modern sampling could reproduce on its own. Modern populations carry the fingerprints of twentieth-century management, the fences, the culls, the relocations between reserves… so diversity estimates measured now can quietly mislead us about what is natural and what is recent damage. The Roosevelt and Rainey specimens predate almost all of that, and they are a baseline collected before modern interventions rearranged the whole lion’s habitat.
By showing that early twentieth-century East African populations were more connected than they are now, the research led by Dr. Campana is a reminder that current translocations that move animals between isolated groups are unlikely to be harmful, and may help rebuild the gene flow that once existed naturally [<strong>7</strong>]. More than that, museum collections assembled for reasons that have nothing to do with genomics, in this case a former president’s hunting expedition, can become some of our most powerful instruments for measuring how much has changed in relatively few years to better inform what to do next about it.</p>

<p> </p>

<h2 id="references">References</h2>
<ol>
  <li><a href="https://www.pnas.org/doi/10.1073/pnas.1500664112">Bauer, H. et al. Lion (<em>Panthera leo</em>) populations are declining rapidly across Africa, except in intensively managed areas. Proc Natl Acad Sci USA. 112, 14894–14899 (2015).</a></li>
  <li><a href="https://www.nature.com/articles/s43247-023-00959-3">Nicholson, S. K. et al. Socio-political and ecological fragility of threatened, free-ranging African lion populations. Commun Earth Environ 4, 302 (2023).</a></li>
  <li><a href="https://www.biorxiv.org/content/10.1101/2025.02.26.640359v1">Armstrong, E. E. et al. President Roosevelt’s lions reveal a century of population fragmentation in Africa’s largest carnivore. bioRxiv 2025.02.26.640359 (2025).</a></li>
  <li><a href="https://link.springer.com/article/10.1007/s10592-004-7729-6">Dubach, J. et al. Molecular genetic variation across the southern and eastern geographic ranges of the African lion, <em>Panthera leo</em>. Conservation Genetics 6, 15–24 (2005).</a></li>
  <li><a href="https://academic.oup.com/mbe/article/38/1/48/5871931">Curry, C. J. et al. Spatiotemporal Genetic Diversity of Lions Reveals the Influence of Habitat Fragmentation across Africa. Mol Biol Evol 38, 48–57 (2021).</a></li>
  <li><a href="https://academic.oup.com/edited-volume/61663/chapter-abstract/553450173">Sunseri, T. The African Rinderpest Panzootic, 1888–1897. Oxford Research Encyclopedia of African History (2018).</a></li>
  <li><a href="https://onlinelibrary.wiley.com/doi/full/10.1111/eva.13318">Bertola, L. D. et al. Genetic guidelines for translocations: Maintaining intraspecific diversity in the lion (<em>Panthera leo</em>). Evol Appl 15, 22–39 (2022).</a></li>
</ol>

<p> </p>

<p>Below, Michael shared with us further details about his profile, career, prospects and future projects:
 </p>

<p><strong>1. Briefly introduce yourself. What is your origin story for how you got into science?</strong> <br />
I am a conservation genomicist at the Smithsonian’s National Zoo and Conservation Biology Institute. My primary areas of expertise include ancient and non-invasive DNA analyses, bioinformatics, and computational genomics. I have wanted to be a scientist since I was a small child fascinated by dinosaurs. While earning my undergraduate degree in palaeontology, I became interested in ancient DNA analyses and my research career developed from there.
 </p>

<p><strong>2.	How and/or why did you start working on this project?</strong> <br />
I am the quintessential “cat person”. In 2014, I was recruited to work on the Roosevelt Resurvey project being led by Kris Helgen and Rob Fleischer. The opportunity to work on big cats fulfilled a life-long research dream.
 </p>

<p><strong>3.	Were there any major challenges in this project? How did you overcome them?</strong> <br />
The first challenge was obtaining sufficient sequence data from the museum specimens for analysis. We started the project well before the advent of cost- effective and time-efficient single-stranded library protocol development. Initially, we were planning on using hybridization capture, but we changed our approach at the urging of Ellie Armstrong (then a PhD student at Stanford University). We upgraded our library preparation protocols over time to accommodate the low-coverage whole genome approach. The second major challenge was limited reference data and genomic parameterization for lions (and big cats in general). We solved the latter problem by generating these data ourselves.
 </p>

<p><strong>4.	What do you think are the main take-home messages of this project?</strong> <br />
Analysis of museum specimens is indispensable for effective conservation actions. The genomic patterns observed using only extant samples often give a very misleading picture of what lion diversity “should” look like. Secondly, analysis of non-model organisms (especially non-humans) is surprisingly challenging due to limited genomic resources, even for iconic species such as lions.
 </p>

<p><strong>5.	What do you think is missing in the field that you would like to work on?</strong> <br />
The translation of ancient-DNA-based conservation recommendations to effective conservation practice remains limited. Implementing effective conservation is much more challenging than the basic science at this point. Part of the issue is that the basic science is still monopolized by the Global North. To address this, I am helping build conservation genomics capacity in range countries.
 </p>

<p><strong>6.	Where do you see yourself in the near future?</strong> <br />
I am planning on continuing my conservation genomics work on a wide range of organisms, including bears, Hawaiian birds, invasive pathogens, dholes, and big cats (among others). One of the best things about working at the zoo is the diversity of projects I get!
 </p>

<p><strong>7.	Free space to tell something you would like to remark.</strong> <br />
I never would have predicted the direction my research career would take, even as a graduate student studying ancient DNA. I advise early-career researchers to keep their aspirations open and be willing to adapt to opportunities as they become available.</p>]]></content><author><name>AaRC</name></author><category term="Blog" /><summary type="html"><![CDATA[July 2026 Tracing lion’s decline through the bones of Roosevelt expeditions “But the lions now offer, and have always offered, the chief source of unpleasant excitement.” by Theodore Roosevelt (African Game Trails, 1910).]]></summary></entry><entry><title type="html">New Steering Committee.</title><link href="https://animal-adna-research-community.github.io/AaRC.github.io//news/2026/05/20/New-Steering-Committee/" rel="alternate" type="text/html" title="New Steering Committee." /><published>2026-05-20T00:00:00+00:00</published><updated>2026-05-20T00:00:00+00:00</updated><id>https://animal-adna-research-community.github.io/AaRC.github.io//news/2026/05/20/New-Steering-Committee</id><content type="html" xml:base="https://animal-adna-research-community.github.io/AaRC.github.io//news/2026/05/20/New-Steering-Committee/"><![CDATA[<p>With AaRC making two years on April 2026, the funding members of the Steering Committee stepped down from their positions and let the new members take charge. 
Pedro Morell Miranda will step down as chair, but remain as webmaster. 
After some delay on our side, <a href="https://animal-adna.org/steering_committee/">the new composition of the Steering Committee is on the website</a>, along with a new section for former members.</p>

<p>We would like to thank all former members for their work and great contributions on making AaRC what it is today.</p>]]></content><author><name>AaRC</name></author><category term="News" /><summary type="html"><![CDATA[With AaRC making two years on April 2026, the funding members of the Steering Committee stepped down from their positions and let the new members take charge. Pedro Morell Miranda will step down as chair, but remain as webmaster. After some delay on our side, the new composition of the Steering Committee is on the website, along with a new section for former members.]]></summary></entry><entry><title type="html">Interview to our February AaRCTikTalks speaker Anne Kathrine Runge</title><link href="https://animal-adna-research-community.github.io/AaRC.github.io//blog/2026/05/01/Interview-February-Speaker2/" rel="alternate" type="text/html" title="Interview to our February AaRCTikTalks speaker Anne Kathrine Runge" /><published>2026-05-01T00:00:00+00:00</published><updated>2026-05-01T00:00:00+00:00</updated><id>https://animal-adna-research-community.github.io/AaRC.github.io//blog/2026/05/01/Interview-February-Speaker2</id><content type="html" xml:base="https://animal-adna-research-community.github.io/AaRC.github.io//blog/2026/05/01/Interview-February-Speaker2/"><![CDATA[<h2 id="february-2026">February 2026</h2>
<h1 id="tracing-prehistoric-zoonoses-through-the-bones-of-ancient-animals">Tracing prehistoric zoonoses through the bones of ancient animals</h1>
<p><em>“Between animal and human medicine there is no dividing line, nor should there be. The object is different but the experience obtained constitutes the basis of all medicine.”</em> (Rudolf Virchow, 1856; by Klauder, J.V., Interrelations of human and veterinary medicine. N Engl J Med, 258:170-177, 1958 [<strong>1</strong>]).</p>

<p> </p>

<h2 id="shared-origins-and-fates">Shared origins and fates</h2>
<p>More than a century and a half ago, Rudolf Virchow, who coined the very term “zoonosis”, recognized that the health of humans and animals is fundamentally intertwined. The words above capture something that remains as relevant today as it was then: that studying disease in one species inevitably teaches us about disease in others. Today, this idea lives on as the One Health concept, which frames infectious disease as a problem that cuts across species boundaries. We are now aware that roughly 60% of known human pathogens are zoonotic, capable of being transmitted between animals and humans <strong>[2]</strong>, and that many of the infectious diseases that have shaped human history, from plague to influenza, jumped from animals to people at some point in the past <strong>[3,4]</strong>. A major question, however, is when and how these zoonotic transitions happened, and whether the shift to farming and animal husbandry during the Neolithic created the conditions for new pathogens to cross into human populations. Ancient DNA has become a powerful tool for investigating these questions in human remains, but the animal side of the equation has remained largely unexplored.
 </p>

<h2 id="bones-that-tell-stories">Bones that tell stories</h2>
<p>Our AaRC speaker Dr. Anne Kathrine Runge described how she and a large team of collaborators set out to tackle a deceptively simple question: can we systematically recover ancient pathogen DNA from animal bones? And if so, can visible signs of disease on those bones help us pick the right specimens to sample? The challenge was considerable. Unlike human burials, where skeletons are often found articulated and relatively well preserved, animal remains in the archaeological record are mostly found as product of consumption or household waste: fragmented, scattered, exposed to the elements, and sometimes heated or boiled during food preparation. All of this inevitably degrades the DNA contained within. On top of that, most animals under human management died from slaughter rather than disease, and sick individuals may have been sacrificed early upon showing signs of infection, further reducing the chances of finding specimens that still carry traces of any pathology and microorganisms associated.
 </p>

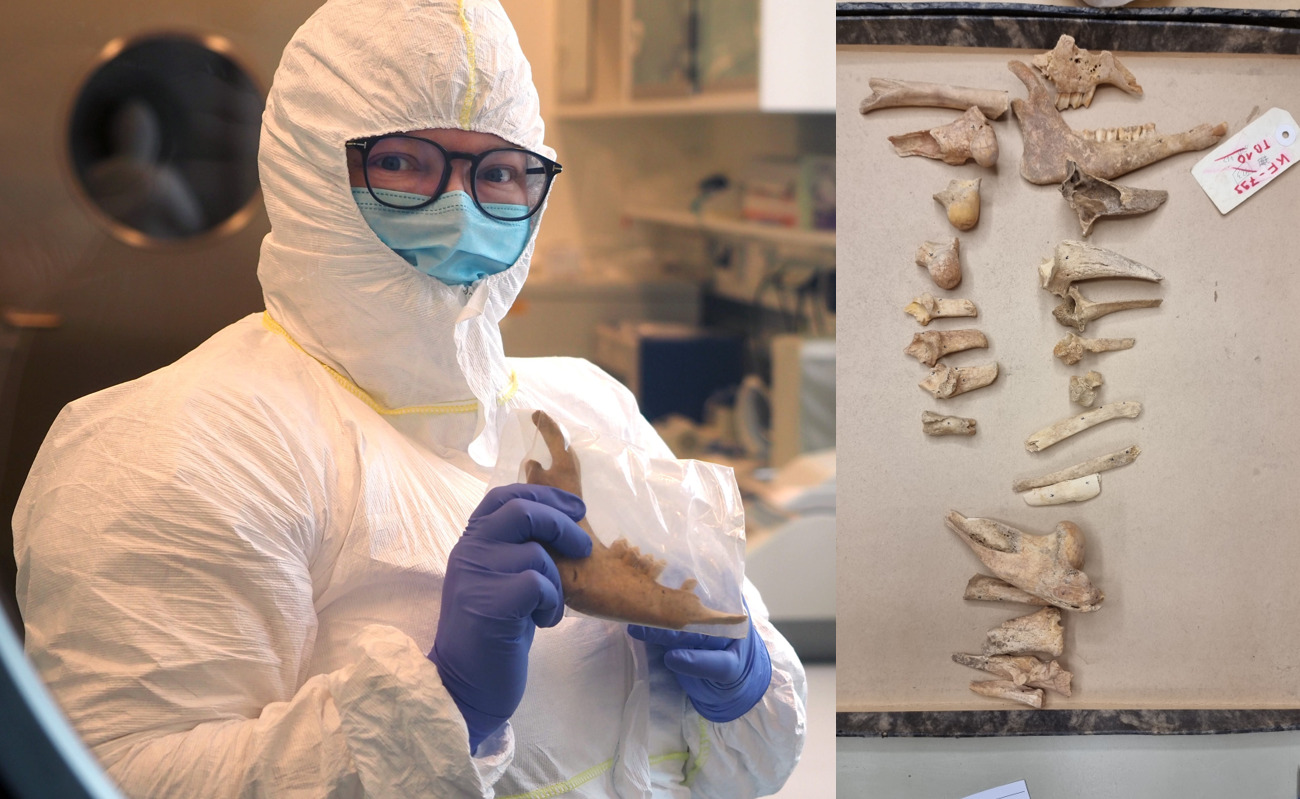
<p>To work around these difficulties, Anne Kathrine and her collaborators assembled a remarkable collection of 346 skeletal elements from at least 328 individual animals, spanning 34 Eurasian sites and roughly six millennia of human history, from the Neolithic to the Medieval period <strong>[5]</strong>. They focused especially on the Bronze Age, a period when human-derived ancient pathogen genomes have been repeatedly recovered in the literature <strong>[6]</strong>. The specimens came mostly from domesticated species (cattle, sheep, pigs, goats, and dogs) but also included wild animals. Critically, about half of the bones were selected because they showed palaeopathological lesions suggestive of infection: periostitis, osteolytic changes, abscesses, or other indicators of potential infectious disease. The rest were teeth or bones without visible lesions, serving as points of comparison.</p>

<p> </p>

<p><img src="/assets/media/Blog_February_2026_AKR_Figure.jpg" alt="ARunge photo" title="ARunge photo" /></p>

<p><em>Dr. Anne Kathrine Runge in the aDNA clean lab holding up a mandible about to be drilled for DNA extraction (Photo credit: Christian Denkhaus). On the right is a selection of pathological bones from the Tilla Bulak site in Uzbekistan (Photo credit: Anne Kathrine Runge).</em></p>
<p> </p>

<h2 id="what-the-screening-revealed">What the screening revealed</h2>
<p>After sequencing, metagenomic screening identified 116 authentic ancient DNA signatures from 29 bacterial species across 55 samples. These included primarily pathogenic species like <em>Salmonella enterica</em>, <em>Erysipelothrix rhusiopathiae</em>, <em>Coxiella burnetii</em>, and <em>Bordetella petrii</em>, alongside a range of opportunistic pathogens from the oral and gastrointestinal microbiome. No viral or eukaryotic parasites were detected.
One of the most telling results was that palaeopathological lesions were significantly enriched for pathogen DNA recovery. Among bones with visible lesions, 23.3% produced robust bacterial hits, compared to 8.4% of teeth and none of the 27 bones without lesions. This supports the idea that expert palaeopathological assessment can meaningfully guide specimen selection for ancient pathogen studies in animals, something that had not been systematically tested before. Interestingly, Dr. Runge also observed striking geographic variation: the Bronze Age site of Tilla Bulak in Uzbekistan yielded a disproportionate number of hits, possibly reflecting better overall DNA preservation at that site, or perhaps a genuinely higher pathogenic pressure during that particular period and location.
 </p>

<h2 id="placing-ancient-pathogens-on-the-evolution-tree">Placing ancient pathogens on the evolution tree</h2>
<p>For two of the recovered species, <em>E. rhusiopathiae</em> and <em>S. lutetiensis</em>, the authors had enough data to attempt phylogenetic placement of the ancient genomes within the known modern diversity. This is far from straightforward with low-coverage shotgun data, so they developed an approach where they first identified variable positions among high-quality modern genomes and then assessed base calls at those positions in the ancient samples. For <em>E. rhusiopathiae</em>, a genome recovered from a Bronze Age cattle tooth excavated at the site of Marinskaya 5 in the North Caucasus (Russia) clustered basally with previously published medieval genomes from Iberian human remains, suggesting that this pathogen was already spreading across multiple hosts in the past. Similarly, three ancient <em>S. lutetiensis</em> genomes from sheep and goat specimens at Tilla Bulak formed a monophyletic group basal to all known modern diversity in the species.
 </p>

<h2 id="challenges-ahead">Challenges ahead</h2>
<p>As Anne Kathrine described, this study represents an important first step toward expanding palaeomicrobiology into the zooarchaeological record. Her results show that pathogen DNA can indeed be recovered from animal remains despite the many challenges involved, and that palaeopathological analysis, which remains rare in animal contexts, can make a real difference in guiding sampling strategies. At the same time, the study highlights several open questions: Lesions are non-specific and cannot point to a particular pathogen, which limits how sampling strategies can be tailored to specific infections. The more porous remodeled bone within lesions may actually work against long-term DNA preservation. And the low coverage obtained from shotgun screening alone is not sufficient for in-depth genomic analyses, meaning that future work using targeted enrichment may be needed to reconstruct more complete pathogen genomes from animal remains.
More broadly, this work brings the One Health perspective into the deep past. Recently, this retrospective approach has been formalized under the concept of One Paleopathology <strong>[7]</strong>, which advocates for integrating archaeological evidence and deep-time perspectives on health across humans, animals, and the environment. By combining palaeopathological analysis, zooarchaeology, and ancient genomics, the work lead by Dr. Runge and her collaborators provides a concrete example of how this framework can be put into practice, opening a path to tracing zoonotic disease reservoirs, spillover events, and pathogen adaptation across millennia.</p>

<p> </p>

<h2 id="references">References</h2>
<ol>
  <li><a href="https://www.nejm.org/doi/full/10.1056/NEJM195801232580405">Klauder, J.V. Interrelations of human and veterinary medicine. N Engl J Med, 258:170-177 (1958).</a></li>
  <li><a href="https://wwwnc.cdc.gov/eid/article/11/12/05-0997_article">Woolhouse, M.E.J. &amp; Gowtage-Sequeria, S. Host range and emerging and reemerging pathogens. Emerg Infect Dis 11: 1842-1847 (2005).</a></li>
  <li><a href="https://www.annualreviews.org/content/journals/10.1146/annurev.ento.50.071803.130337">Gage, K. L. &amp; Kosoy, M. Y. Natural history of plague: perspectives from more than a century of research. Annu Rev Entomol 50, 505–528 (2005).</a></li>
  <li><a href="https://pmc.ncbi.nlm.nih.gov/articles/PMC2720273/">Taubenberger, J. K. The Origin and Virulence of the 1918 “Spanish” Influenza Virus. Proc Am Philos Soc 150, 86 (2006).</a></li>
  <li><a href="https://www.nature.com/articles/s41467-026-71543-4">Runge, A.K.W. et al. Probing the zooarchaeological record across time and space for ancient pathogen DNA. Nat Commun 17, 3469 (2026).</a></li>
  <li><a href="https://www.nature.com/articles/s41586-025-09192-8">Sikora, M. et al. The spatiotemporal distribution of human pathogens in ancient Eurasia. Nature 643, 1011–1019 (2025).</a></li>
  <li><a href="https://academic.oup.com/bioscience/advance-article-abstract/doi/10.1093/biosci/biaf115/8198379?login=false">Buikstra, J.E., Uhl, E.W. &amp; Robbins Schug, G. One Paleopathology and lessons from the past. BioScience, biaf115 (2025).</a></li>
</ol>

<p> </p>

<p>Below, Anne Kathrine shared with us further details about her profile, career, prospects and future projects:
 </p>

<p><strong>1. Briefly introduce yourself. What is your origin story for how you got into science?</strong> <br />
I grew up in the Danish countryside with nature just outside my door. One of my grandfathers was a farmer who loved animals and plants, and the other was so enthused by beetles that he was one of the leading entomologists in the country, and I loved following them around learning about all the different species that surround us. For this reason, no one was really surprised that I wanted to study biology. At uni I fell in love with genetics then genomics, and especially ancient genomics, and I was lucky that I got the opportunity to pursue this interest.
 </p>

<p><strong>2. How and/or why did you start working on this project?</strong> <br />
That was a bit of a happy accident, I think. Originally the project was quite different, but despite having everything in place and verbally agreed on, in the end we did not get the sampling permit. So we had to pivot, and fortunately our amazing collaborators stepped in with samples from sites they had already worked on. Together we made it work, and I’m really grateful for everyone who contributed to that.
 </p>

<p><strong>3. Were there any major challenges in this project? How did you overcome them?</strong> <br />
I think the biggest challenge in the project was that we could not know beforehand which pathogen we would find if we managed to get DNA at all. There are a lot of really interesting questions that can be asked about zoonotic diseases, early spillover events, reservoir populations, host adaptation, and evolution, but they hinge on being able to obtain specific pathogen DNA from non-specific lesions that are less than optimal for DNA preservation. We overcame this by processing a lot of samples, but even then, we only had a handful of samples that produced something really interesting, and for these, we still only had the screening data and could not do anything in depth.
 </p>

<p><strong>4. What do you think are the main take-home messages of this project?</strong> <br />
Genuinely, I think the take-home message is that this was really difficult, but also that there are ways to make studying ancient pathogens from animal remains easier. It does require collaboration with an expert in palaeopathology, and these people, especially those that focus on animal pathology, are rare. I also think that there are some really interesting implications for future studies and I hope it will help future researchers design some super cool studies.
 </p>

<p><strong>5. What do you think is missing in the field that you would like to work on?</strong> <br />
The biggest thing? That’s definitely the ability to look at a lesion and know which pathogen caused it. Unfortunately, that is unlikely to become any easier and looking for specific pathogens in animal remains is likely going to continue to be really difficult. What can be worked on, however, is sampling strategies. For example, where on the bone is the best place to sample, as well as optimizing DNA extraction methods for pathogen DNA.
 </p>

<p><strong>6. Where do you see yourself in the near future?</strong> <br />
I see myself back in Denmark and hopefully in an industry job. I have not quite figured out what I want, but I know it is not academia.
 </p>

<p><strong>7. Free space to tell something you would like to remark.</strong> <br />
I’m really looking forward to seeing where this field will go in the future.</p>]]></content><author><name>AaRC</name></author><category term="Blog" /><summary type="html"><![CDATA[February 2026 Tracing prehistoric zoonoses through the bones of ancient animals “Between animal and human medicine there is no dividing line, nor should there be. The object is different but the experience obtained constitutes the basis of all medicine.” (Rudolf Virchow, 1856; by Klauder, J.V., Interrelations of human and veterinary medicine. N Engl J Med, 258:170-177, 1958 [1]).]]></summary></entry><entry><title type="html">Interview to our February AaRCTikTalks speaker Anna Nagel</title><link href="https://animal-adna-research-community.github.io/AaRC.github.io//blog/2026/04/20/Interview-February-Speaker1/" rel="alternate" type="text/html" title="Interview to our February AaRCTikTalks speaker Anna Nagel" /><published>2026-04-20T00:00:00+00:00</published><updated>2026-04-20T00:00:00+00:00</updated><id>https://animal-adna-research-community.github.io/AaRC.github.io//blog/2026/04/20/Interview-February-Speaker1</id><content type="html" xml:base="https://animal-adna-research-community.github.io/AaRC.github.io//blog/2026/04/20/Interview-February-Speaker1/"><![CDATA[<h2 id="february-2026">February 2026</h2>
<h1 id="dating-ancient-dna-under-the-multispecies-coalescent">Dating ancient DNA under the multispecies coalescent</h1>
<p><em>“The affinities of all the beings of the same class have sometimes been represented by a great tree. I believe this simile largely speaks the truth. The green and budding twigs may represent existing species; and those produced during each former year may represent the long succession of extinct species.”</em> (Charles Darwin, On the Origin of Species by Means of Natural Selection, Chapter IV, 1859).</p>

<p> </p>

<h2 id="the-problem-of-time">The problem of time</h2>
<p>Estimating when species diverged is one of the central goals of evolutionary biology. Since molecular sequences accumulate mutations over time, by comparing DNA from different species we can infer how long ago they shared a common ancestor. But there is an inherent limitation: molecular data alone only tell us about relative divergence, not absolute time. This means that without some external reference point, we cannot separate the speed of molecular evolution from the duration over which it has occurred. Researchers have traditionally relied on fossil age calibrations to link phylogenetic trees to an absolute timescale. However, placing fossils on the right branches of a phylogeny is difficult, since they must rely on morphological characters that may be sparse or ambiguous.
Ancient DNA (aDNA) offers a different approach. Because aDNA samples come with known ages, from radiocarbon dating or stratigraphic context, they effectively provide timestamps scattered across the branches of a phylogeny. If the time intervals between samples are large enough, or the number of loci is large enough, the expected difference in accumulated mutations between lineages sampled at different times becomes detectable, making it possible to estimate mutation rates and divergence times directly from the data.
 </p>

<h2 id="when-gene-trees-are-not-species-trees">When gene trees are not species trees</h2>
<p>Our AaRC speaker Dr. Anna Nagel described how she developed a new framework to bring tip dating into the multispecies coalescent (MSC) model, implemented in the widely used program BPP [<strong>1</strong>]. Why does this matter? Most existing methods that incorporate sample ages into phylogenetic dating, such as BEAST, estimate gene trees rather than species trees. As Anna mentioned, this distinction is critical, since <em>“the common ancestor of a gene must be older than the common ancestor of the species”</em> [<strong>1</strong>]. As a result, using gene tree divergence times as proxies for species divergence times leads to systematic overestimation. The topologies of different gene trees may also vary, and the coalescence times will almost certainly differ for each locus. The MSC explicitly models gene trees within species trees, accounting for ancestral population sizes, and produces unbiased estimates of species-level divergence times.
While the MSC has been available in BPP for years, it previously assumed all sequences were sampled at the present. Anna and her collaborators extended the model to accommodate sequences sampled at different times within each population [<strong>1</strong>].</p>

<p> </p>

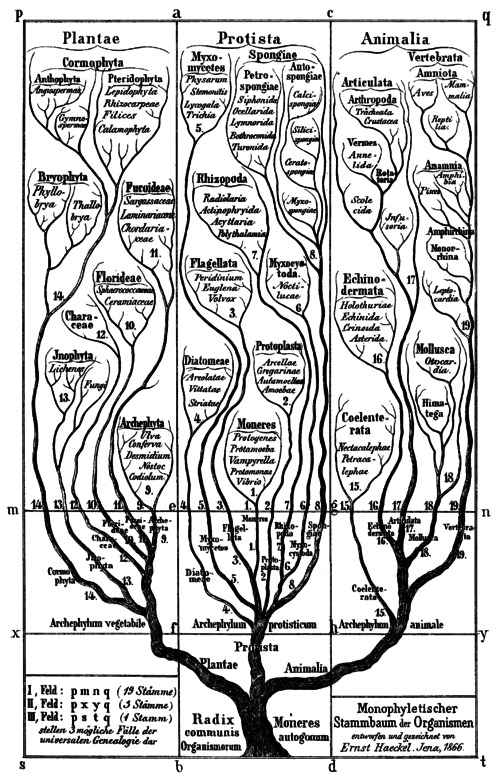
<p><img src="/assets/media/Blog_February_2026_AN_Figure.jpg" alt="Evolution tree" title="Evolution tree" /></p>

<p><em>"Monophyletischer Stammbaum der Organismen" (Monophyletic Tree of Organisms). From Ernst Haeckel, Generelle Morphologie der Organismen, Vol. II (Georg Reimer, Berlin, 1866). Via Wikimedia Commons.</em></p>
<p> </p>

<h2 id="what-the-simulations-revealed">What the simulations revealed</h2>
<p>Dr. Nagel tested the method extensively through simulations under a range of biologically realistic scenarios. These included populations with divergence times spanning from a few thousand to millions of years. The results were encouraging: with sufficient data, particularly many loci, the method accurately recovered both mutation rates and divergence times. The simulations also showed that the precision of estimates improved most when prioritizing two factors: increasing the number of loci and using older ancient samples. Perhaps unsurprisingly, older samples provide a longer temporal span, making the difference in accumulated mutations more detectable across lineages.
One of the most informative parts of the study explored what happens when researchers ignore sample ages and treat all aDNA as contemporary. The consequences were substantial: divergence times were biased, the mutation rate became unidentifiable, and effective population sizes for extinct species were overestimated. These findings serve as a cautionary note for the field, and suggest that even with relatively young aDNA samples, failing to model sample ages properly can lead to misleading inferences.
 </p>

<h2 id="from-mammoths-to-elephants">From mammoths to elephants</h2>
<p>To put the method to work on real data, Dr. Nagel analyzed two genomic datasets of elephants and mammoths. One dataset consisted of mitochondrial genomes, including ancient woolly mammoth sequences [<strong>2</strong>], and the other was a nuclear dataset with sequences from Asian, forest, and savanna elephants, woolly mammoths, and American mastodons [<strong>3</strong>]. The mitochondrial analysis produced divergence time estimates that were substantially younger than previous estimates, with DNA deamination damage possibly driving inflated mutation rates. This finding highlights a key challenge that the aDNA field still faces: standard damage-filtering approaches may not fully remove the effects of post-mortem modifications, and future work may benefit from explicitly modeling DNA damage itself, as the authors discuss [<strong>1</strong>].
The nuclear dataset, despite having short loci and limited data, yielded divergence estimates broadly consistent with previous studies that relied on fossil-based calibration. However, the simulations clearly showed that as datasets grow in size, the impact of incorrect sample dates increases, making proper tip dating a relevant issue in the era of large-scale paleogenomic studies.
 </p>

<h2 id="looking-forward">Looking forward</h2>
<p>Dr. Anna Nagel’s work opens several avenues for future development. A practical challenge that remains is the method’s convergence, which the authors found became harder as dataset size increased. Developing better strategies will be important to scale to larger genomic datasets including more loci. The current implementation assumes a fixed species tree topology and no gene flow, assumptions that the latest versions of BPP can relax for contemporary data but not yet for tip-dated analyses. Incorporating migration models would be particularly valuable for systems such as elephants, where hybridization between lineages is well documented, or for hominin datasets where gene flow between archaic and modern populations is a central question. Additionally, combining tip dating with fossil calibrations, and developing models that explicitly account for DNA degradation, could further improve the accuracy and utility of this framework.</p>

<p> </p>

<h2 id="references">References</h2>
<ol>
  <li><a href="https://academic.oup.com/sysbio/article/73/6/964/7724094?login=false">Nagel, A.A. et al. Bayesian inference under the multispecies coalescent with ancient DNA sequences. Syst Biol 73: 964–978 (2024).</a></li>
  <li><a href="https://www.nature.com/articles/s41586-021-03224-9">van der Valk, T. et al. Million-year-old DNA sheds light on the genomic history of mammoths. Nature 591: 265–269 (2021).</a></li>
  <li><a href="https://journals.plos.org/plosbiology/article?id=10.1371/journal.pbio.1000564">Rohland, N. et al. Genomic DNA sequences from mastodon and woolly mammoth reveal deep speciation of forest and savanna elephants. PLoS Biol 8: e1000564 (2010).</a></li>
</ol>

<p> </p>

<p>Below, Anna shared with us further details about her profile, career, prospects and future projects:
 </p>

<p><strong>1. Briefly introduce yourself. What is your origin story for how you got into science?</strong> <br />
I am a postdoc Washington University in St. Louis. I work on population genetics and phylogenetics methods development, mostly relating to the estimation of times. I was always excited about science, but I started as a chemistry major. I was interested in Ecology from spending time at an environmental learning center as a kid, but I thought biology wasn’t mathematical enough. After taking Evolution, I realized that wasn’t true and mathematical models are a powerful tool for applications ranging from proof of concept to inference. Moreover, I was really excited by how studying evolutionary theory allows us to understand the patterns of biological diversity across scales, from individual loci to the tree of life.
 </p>

<p><strong>2. How and/or why did you start working on this project?</strong> <br />
I was actually thinking about HIV transmission and how coalescent models could be used to model transmission with multiple sequences from each infected individual. That project would have required a lot of additions to existing software, including adding tip dating to the multispecies coalescent (MSC). I decided to work on aDNA first because it was a much smaller project. I still haven’t gotten back to the HIV transmission project.
 </p>

<p><strong>3. Were there any major challenges in this project? How did you overcome them?</strong> <br />
One of the main obstacles was getting the MCMCs to mix. Addressing this required developing MCMC proposals specific to tip dating.
 </p>

<p><strong>4. What do you think are the main take-home messages of this project?</strong> <br />
Even relatively young age sequences (in comparison to the species divergence times) can give a lot of information about species divergence times and allow us to estimate those times without outside information such as fossil ages. However, DNA degradation may be a pervasive problem in these types of analyses and future work should explicitly model DNA degradation.
 </p>

<p><strong>5. What do you think is missing in the field that you would like to work on?</strong> <br />
I would be excited to work on migration models with aDNA. Bayesian inference with coalescent models with migrations are difficult to get to work correctly because they are really complex. However, these models would allow us to analyze a more datasets more satisfactorily. In my empirical analysis, I removed a mammoth species that had been suggested to have been a product of hybrid speciation, and likely my estimate of divergence time estimate of the African elephant species was biased substantially due to hybridization. Hominid datasets would also require migration models.
 </p>

<p><strong>6. Where do you see yourself in the near future?</strong> <br />
Right now, I’m hoping to keep working on similar types of questions relating to Bayesian phylogenetic inference and the intersection of phylogenetics and population genetics. I’ve also been working on machine learning in phylogenetics, so I’d like to see how those methods could be applied to the MSC.</p>]]></content><author><name>AaRC</name></author><category term="Blog" /><summary type="html"><![CDATA[February 2026 Dating ancient DNA under the multispecies coalescent “The affinities of all the beings of the same class have sometimes been represented by a great tree. I believe this simile largely speaks the truth. The green and budding twigs may represent existing species; and those produced during each former year may represent the long succession of extinct species.” (Charles Darwin, On the Origin of Species by Means of Natural Selection, Chapter IV, 1859).]]></summary></entry><entry><title type="html">AaRCademy - Relatedness</title><link href="https://animal-adna-research-community.github.io/AaRC.github.io//events/2026/04/16/AaRCademy-Relatedness/" rel="alternate" type="text/html" title="AaRCademy - Relatedness" /><published>2026-04-16T00:00:00+00:00</published><updated>2026-04-16T00:00:00+00:00</updated><id>https://animal-adna-research-community.github.io/AaRC.github.io//events/2026/04/16/AaRCademy-Relatedness</id><content type="html" xml:base="https://animal-adna-research-community.github.io/AaRC.github.io//events/2026/04/16/AaRCademy-Relatedness/"><![CDATA[<p><img src="/assets/media/AaRCAdemy_April16.png" alt="AaRCademy Relatedness poster" title="April 16 workshop" /></p>

<p>One of the coolest ways to use ancient DNA to explore the past is to try to reconstruct the relationship between individuals and populations in the past.
Relatedness methods are quite common in human studies, where a lot of times you know from the start that several samples come from the same period and region, so 
we can explore if they were buried together because they were related “by blood” or if something else happened. In animals, however, this approach is less common. 
Shorter generation times, less careful burials and for most species a low density of samples make direct relatedness dificult to test. That, however, doesn’t mean we can’t use some of these methods to 
study how our populations are related to one another. With that in mind, come and join us and Jolijn Erven (University College Dublin) on <em>April 16th, 14:00 CET</em> to learn more about all the ways we can explore 
palaeogenomic data to describe the relatedness between our samples.</p>

<p>To register, follow the QR code or click <a href="https://docs.google.com/forms/d/1kUg0hMJOK-b3Y8q2mjXS3uoeKZMEIxrg0QiJNpMNIzM/edit">here</a></p>]]></content><author><name>AaRC</name></author><category term="Events" /><category term="AaRCademy," /><category term="Workshops" /><summary type="html"><![CDATA[]]></summary></entry><entry><title type="html">AaRCTikTalks March 2026</title><link href="https://animal-adna-research-community.github.io/AaRC.github.io//events/2026/03/27/MarchAaRCTikTalks/" rel="alternate" type="text/html" title="AaRCTikTalks March 2026" /><published>2026-03-27T00:00:00+00:00</published><updated>2026-03-27T00:00:00+00:00</updated><id>https://animal-adna-research-community.github.io/AaRC.github.io//events/2026/03/27/MarchAaRCTikTalks</id><content type="html" xml:base="https://animal-adna-research-community.github.io/AaRC.github.io//events/2026/03/27/MarchAaRCTikTalks/"><![CDATA[<p><img src="/assets/media/aarc_talks_mar2026.png" alt="AaRCTikTalks poster for March 2026" title="AaRCTikTalks poster for March 2026" /></p>

<p>For March the AaRCTikTalks go to the Land Down Under! We’ll have a matinée (in CET time) so we can host two amazing talks focused on Australia:</p>

<ul>
  <li>Loukas Koungoulos: Origins and History of Australia’s Native Wild Dog</li>
  <li>Siobhan Evans: Tracing the Fate of Ancient DNA in Australian Cave Sediments</li>
</ul>

<p>As with previous AaRCTikTalks, you can register to the seminar mailing list <a href="https://docs.google.com/forms/d/e/1FAIpQLSfq4BUmArgA96iWjm71ocQwbosLZRNb72rZK8Oky1RCnJsGNw/viewform">here</a>
or join us on <a href="https://matrix.to/#/#aarc:archaeo.social">Element</a>, where a link will be shared minutes before it starts.</p>]]></content><author><name>AaRC</name></author><category term="Events" /><category term="AaRCTikTalks" /><summary type="html"><![CDATA[]]></summary></entry><entry><title type="html">AaRCTikTalks January 2026</title><link href="https://animal-adna-research-community.github.io/AaRC.github.io//events/2026/01/30/JanuaryAaRCTikTalks/" rel="alternate" type="text/html" title="AaRCTikTalks January 2026" /><published>2026-01-30T00:00:00+00:00</published><updated>2026-01-30T00:00:00+00:00</updated><id>https://animal-adna-research-community.github.io/AaRC.github.io//events/2026/01/30/JanuaryAaRCTikTalks</id><content type="html" xml:base="https://animal-adna-research-community.github.io/AaRC.github.io//events/2026/01/30/JanuaryAaRCTikTalks/"><![CDATA[<p><img src="/assets/media/aarc_talks_jan2026.png" alt="AaRCTikTalks poster for January 2026" title="AaRCTikTalks poster for September" /></p>

<p>New year, new AaRC TikTalks! And we start strong with 2 amazing presenters:</p>

<ul>
  <li>Tatiana Feuerborn: Tracind the Legacy of Greenland  Sled Dogs throught time.</li>
  <li>Anianna Wingarten: Mitogenomes of Middle Pleistocene horses from Schöningen.</li>
</ul>

<p>As with previous AaRCTikTalks, you can register to the seminar mailing list <a href="https://docs.google.com/forms/d/e/1FAIpQLSfq4BUmArgA96iWjm71ocQwbosLZRNb72rZK8Oky1RCnJsGNw/viewform">here</a>
or join us on <a href="https://matrix.to/#/#aarc:archaeo.social">Element</a>, where a link will be shared minutes before it starts.</p>]]></content><author><name>AaRC</name></author><category term="Events" /><category term="AaRCTikTalks" /><summary type="html"><![CDATA[]]></summary></entry><entry><title type="html">Interview to our January AaRCTikTalks speaker Arianna Weingarten</title><link href="https://animal-adna-research-community.github.io/AaRC.github.io//blog/2026/01/12/Interview-January-Speaker1/" rel="alternate" type="text/html" title="Interview to our January AaRCTikTalks speaker Arianna Weingarten" /><published>2026-01-12T00:00:00+00:00</published><updated>2026-01-12T00:00:00+00:00</updated><id>https://animal-adna-research-community.github.io/AaRC.github.io//blog/2026/01/12/Interview-January-Speaker1</id><content type="html" xml:base="https://animal-adna-research-community.github.io/AaRC.github.io//blog/2026/01/12/Interview-January-Speaker1/"><![CDATA[<h2 id="january-2026">January 2026</h2>
<h1 id="ancient-dna-from-open-air-sites-reveals-new-insights-about-horse-evolution">Ancient DNA from open-air sites reveals new insights about horse evolution</h1>
<p><em>“Do you give the horse his strength? Do you clothe his neck with a mane? Do you make him leap like the locust? His majestic snorting is terrifying. He paws in the valley and exults in his strength; he goes out to meet the weapons. He laughs at fear and is not dismayed; he does not turn back from the sword.”</em> (From the Book of Job 39:19-25).</p>

<p> </p>

<h2 id="the-horse-legacy">The horse legacy</h2>
<p>For millennia, humans have celebrated the horse’s power, grace, and fearlessness. The text above captures something profound about these magnificent animals, their strength, their courage, their deep connection to human history. While the book of Job marvels at the power of the horse, long before horses carried riders or pulled chariots, their ancestors were already roaming prehistoric landscapes. The horse family (<em>Equidae</em>) has a fossil history stretching back about 55 million years, making it a key example for studying long-term evolution. Today, only one genus survives, Equus, which includes zebras, donkeys, and horses. However, fossils show that the family was once far more diverse, with more than 35 genera and many species [<strong>1</strong>]. Although some earlier lineages survived for a long time, most disappeared. As a result, all modern living horses descend from a single Eurasian lineage that originated from this later migration [<strong>2</strong>].
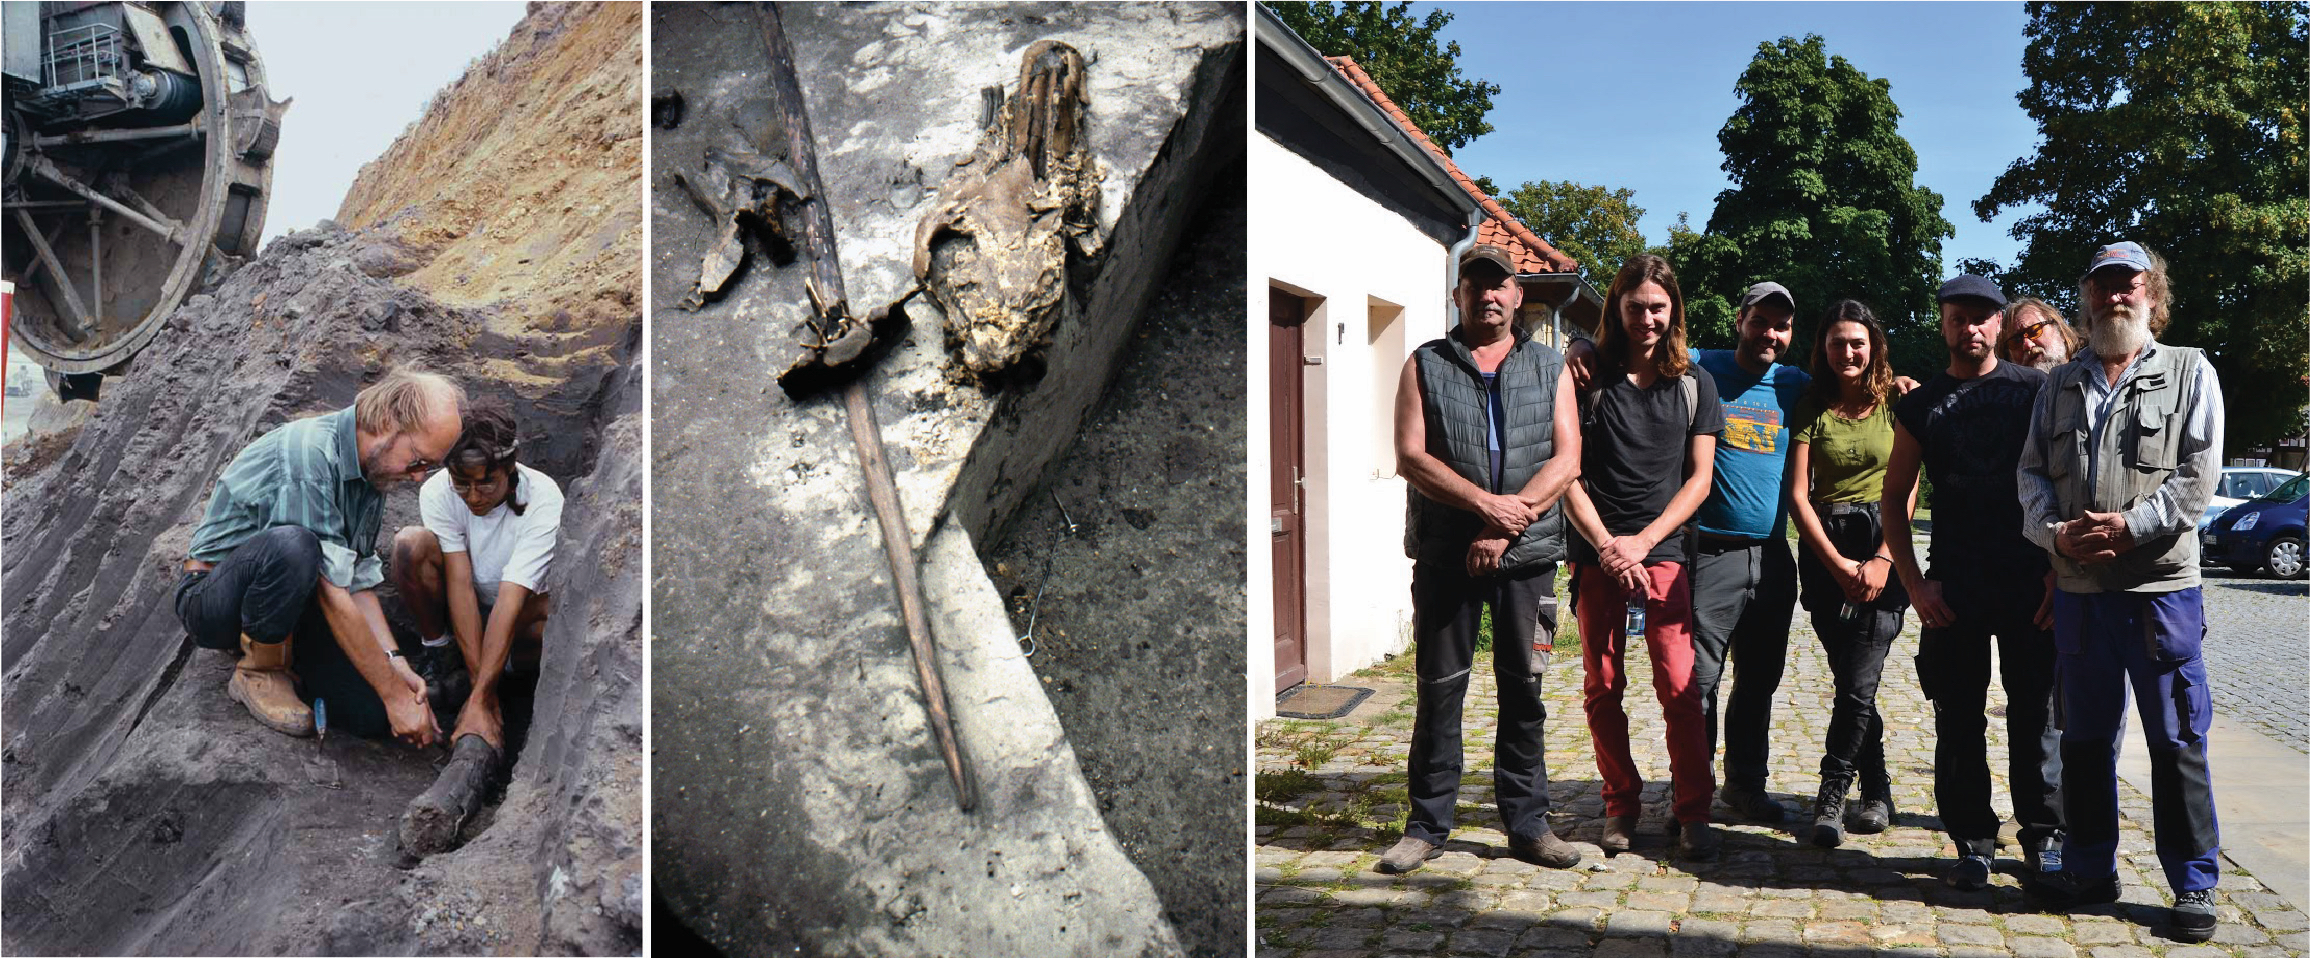
Our AaRC speaker Arianna Weingarten described how an early visit to a remarkable archeological site shaped her interest for paleogenomics research. Located between the cities of Hannover and Berlin, the place was the Middle Pleistocene open-air site complex of Schöningen in Lower Saxony, Germany, dated to about 320–300 thousand years ago. The site itself tells an extraordinary story: Schöningen is where archaeologists discovered the world’s oldest complete wooden spears, found alongside the butchered remains of 20-25 horses [<strong>3,4</strong>]. These artifacts provide direct evidence that our ancient hominin ancestors were already sophisticated hunters with complex tools, and horses were central to their survival.</p>

<p> </p>

<p><img src="/assets/media/Blog_January_2026_AW_Figure.jpg" alt="Schöningen excavation" title="Schöningen excavation" /></p>

<p><em>Left: Dr. Hartmut Thieme uncovering an elephant tusk during a rescue excavation (Photo: Peter Pfarr). Middle: A horse skull discovered next to a wooden spear (Photo: Nicholas J. Conard).  Right: Arianna during her first excavation at Schöningen with the excavation team and researchers Ivo Verheijen and Gabriele Russo (Photo: Jordi Serangeli).</em></p>
<p> </p>

<h2 id="pushing-the-limits-of-ancient-dna">Pushing the limits of ancient DNA</h2>
<p>In their study, Arianna and her collaborators focused on two horse specimens recovered from the site and identified morphologically as Equus mosbachensis [<strong>5</strong>]. Using petrous bones, which often preserve DNA better than other tissues and bone types, they sequenced their mitochondrial genomes and determined that one individual was male and the other female. Extracting DNA from these Middle Pleistocene remains was particularly challenging. As Arianna described, the genetic material was highly fragmented and chemically damaged. To address this, the authors developed a specific computational method called DORIAN to help detect and down-weight damaged DNA bases [<strong>5</strong>], thus improving the accuracy of genome reconstruction. The recovery of such ancient mitochondrial genomes extended the known limit of DNA preservation in open-air sites to roughly 300,000 years, demonstrating that favorable environmental conditions can sometimes preserve genetic material almost as well as caves.
 </p>

<h2 id="reconstructing-the-unbroken-horse-lineage">Reconstructing the unbroken horse lineage</h2>
<p>After such a success, Arianna compared them with more than a hundred ancient and modern horse mitochondrial sequences to assess their evolutionary relationship among past horse lineages. Previous research had established an ancestral clade (B) evolved in North America, which eventually migrated to Eurasia giving rise to two clades (A and C), one of which remigrated to North America during the Late Pleistocene and diversified into two additional clades (A1 and A2) [<strong>6</strong>]. The large megafaunal extinction of the Early Holocene, however, eventually resulted in the extinction of all these except for clade A, which gave rise to all modern horses [<strong>6</strong>]. Phylogenetic analyses carried out in this study showed that the two Schöningen horses occupy a basal position relative to the diversification of modern horses within clade A [<strong>5</strong>]. In simple terms, their maternal lineages split off very early from the clade that eventually led to living horses. Arianna and her collaborators also used molecular dating of the mitochondrial genomes, and determined the age of one of them to be around 360,000-years-old [<strong>5</strong>]. Moreover, they even estimated that the ancestral mitochondrial clade B diverged around 800,000 years ago from A and C clades, and that the A clade diverged from their ancient Schöningen horses around 570,000 years ago. The authors further report that a major diversification within extant clade A likely began after roughly 230,000 years ago, a period overlapping with an interglacial phase that may have influenced equine evolution through environmental change [<strong>5</strong>].
 </p>

<h2 id="new-tools-for-new-challenges">New tools for new challenges</h2>
<p>As Arianna finally highlights, their study shows how technological advances tailored for deep time remains are transforming the palaeogenetics field. Their innovative methods helped reducing bias and recovering more usable data from highly degraded samples. This study not only extends the timeline of what is possible in DNA recovery from open-air sites but also provides crucial data points for reconstructing horse evolution during a period that left few genetic traces.</p>

<p> </p>

<h2 id="references">References</h2>
<ol>
  <li><a href="https://www.nature.com/articles/336466a0">MacFadden, B.J. &amp; Hulbert, R.C. Explosive speciation at the base of the adaptive radiation of Miocene grazing horses. Nature 336: 466–468 (1988).</a></li>
  <li><a href="https://www.frontiersin.org/journals/ecology-and-evolution/articles/10.3389/fevo.2019.00278/full">Rook, L. et al. Mammal biochronology (Land Mammal Ages) Around the world from Late Miocene to Middle Pleistocene and major events in horse evolutionary history. Front Ecol Evol 7: 278 (2019).</a></li>
  <li><a href="https://www.sciencedirect.com/science/article/pii/S0047248424000988">Hutson, J.M. et al. Persistent predators: Zooarchaeological evidence for specialized horse hunting at Schöningen 13II-4. J Hum Evol 196: 103590 (2024).</a></li>
  <li><a href="https://www.science.org/doi/full/10.1126/sciadv.adv0752">Hutson, J.M. et al. Revised age for Schöningen hunting spears indicates intensification of Neanderthal cooperative behavior around 200,000 years ago. Sci Adv 11: eadv0752 (2025).</a></li>
  <li><a href="https://www.nature.com/articles/s41559-025-02859-5">Weingarten, A. et al. Mitochondrial genomes of Middle Pleistocene horses from the open-air site complex of Schöningen. Nat Ecol Evol 9: 2248 (2025).</a></li>
  <li><a href="https://onlinelibrary.wiley.com/doi/10.1111/mec.15977">Vershinina, A.O. et al. Ancient horse genomes reveal the timing and extent of dispersals across the Bering Land Bridge. Mol Ecol 30: 6144–6161 (2021).</a></li>
</ol>

<p> </p>

<p>Below, Arianna shared with us further details about her profile, career, prospects and future projects:
 </p>

<p><strong>1. Briefly introduce yourself. What is your origin story for how you got into science?</strong> <br />
I completed my undergraduate studies at the University of California, Santa Cruz without arriving with a plan to become a scientist. Instead of following a lifelong passion or being pushed to choose a subject early on, I was fortunate to have the freedom to explore different disciplines and gradually discover what I was interested in. I kept taking biology classes because it seemed like important knowledge for navigating the world, while geology classes drew me in with their field trips around California. This mix of practicality and curiosity eventually led me to a double major in biology and a combined anthropology/earth science degree. Somewhere along the way, I was introduced to ancient DNA and was immediately fascinated because it combined my interests of deep-time, geology, archaeology, and biology. This led me to my current position as a PhD student in the Archaeo-and Palaeogenetics group at the University of Tübingen.
 </p>

<p><strong>2.	How and/or why did you start working on this project?</strong> <br />
I actually started this project during my master’s degree, after I went on excavation at Schöningen. At the time there had been a few unpublished attempts at retrieving ancient DNA from the site. The project originated as my master’s thesis, in which we applied the most up-to-date sampling and laboratory protocols, and surprisingly, got some promising results! Building off this initial shotgun screening, my PhD research expanded the work by producing mitochondrial baits in-house and performing mitochondrial capture experiments to investigate a few species from the site. Though only the horse story is out at the moment!
 </p>

<p><strong>3.	Were there any major challenges in this project? How did you overcome them?</strong> <br />
Definitely. One of the major challenges was deciding how to best move forward with the data we generated. The extremely high damage in the samples, paired with the very short fragment lengths, meant that decisions about cut-off thresholds and how to handle damage had a large impact on the reconstruction of the mitogenomes. To deal with this we carried out extensive testing using different software tools and custom scripts. Ultimately, we established a collaboration with bioinformaticians at the University of Tübingen to develop a tool for our data (and similar data types!), which allowed us to keep the maximum amount of reliable information in our downstream analysis for molecular dating and phylogenetic work.
 </p>

<p><strong>4.	What do you think are the main take-home messages of this project?</strong> <br />
The main take-home message of the project is the opportunity for discovery that comes from pushing the boundaries of how far back in time, and across which environmental contexts, genetic data can be retrieved from the fossil record. However, this effort definitely requires keeping in mind a number of caveats, including rigorous data screening to ensure that the reconstructed genomes reflect true biological variation. Overall, it’s a little bit high risk, high reward but that’s also what’s exciting.
 </p>

<p><strong>5.	What do you think is missing in the field that you would like to work on?</strong> <br />
I’m really interested in the exploratory side of the field, so working with previously unknown genomes and stitching together paleontological, archeological, and genetic information. There are still so many extinct species that remain genetically unexplored. Expanding ancient genomic data, especially on the whole-genome-level, can provide a view into past adaptations and population dynamics and I would like to contribute to developing and applying these approaches.
 </p>

<p><strong>6.	Where do you see yourself in the near future?</strong> <br />
Since I’m at the end of my PhD, the near future looks like an intense final period, a hopefully not too horrible ending, followed by a dreamy vacation, and then on to the next adventure!
 </p>

<p><strong>7.	Free space to tell something you would like to remark.</strong> <br />
Thanks again for the invite :)</p>]]></content><author><name>AaRC</name></author><category term="Blog" /><summary type="html"><![CDATA[January 2026 Ancient DNA from open-air sites reveals new insights about horse evolution “Do you give the horse his strength? Do you clothe his neck with a mane? Do you make him leap like the locust? His majestic snorting is terrifying. He paws in the valley and exults in his strength; he goes out to meet the weapons. He laughs at fear and is not dismayed; he does not turn back from the sword.” (From the Book of Job 39:19-25).]]></summary></entry><entry><title type="html">Interview to our January AaRCTikTalks speaker Tatiana Feuerborn</title><link href="https://animal-adna-research-community.github.io/AaRC.github.io//blog/2026/01/12/Interview-January-Speaker2/" rel="alternate" type="text/html" title="Interview to our January AaRCTikTalks speaker Tatiana Feuerborn" /><published>2026-01-12T00:00:00+00:00</published><updated>2026-01-12T00:00:00+00:00</updated><id>https://animal-adna-research-community.github.io/AaRC.github.io//blog/2026/01/12/Interview-January-Speaker2</id><content type="html" xml:base="https://animal-adna-research-community.github.io/AaRC.github.io//blog/2026/01/12/Interview-January-Speaker2/"><![CDATA[<h2 id="january-2026">January 2026</h2>
<h1 id="a-genetic-journey-through-the-ancient-lineage-of-greenlands-qimmit">A genetic journey through the ancient lineage of Greenland’s Qimmit</h1>

<p><em>“The dogs astutely sense that work is at hand. They shake off the new fallen snow, jumping up and down in their eagerness to get going. With assuredness, the musher selects his team of dogs, saving the lead dog, which today happens to be a bitch in golden colors, until the very last minute… Off we go. With a start, you are yanked back on the sled, and some time passes before you can gain your equilibrium. The silence, the enormous expanses of land, the bond between the musher and his dogs and the cold will quickly transport you to another world…”</em> (Anonymous, from <a href="https://visitgreenland.com/">https://visitgreenland.com/</a>)</p>

<p> </p>

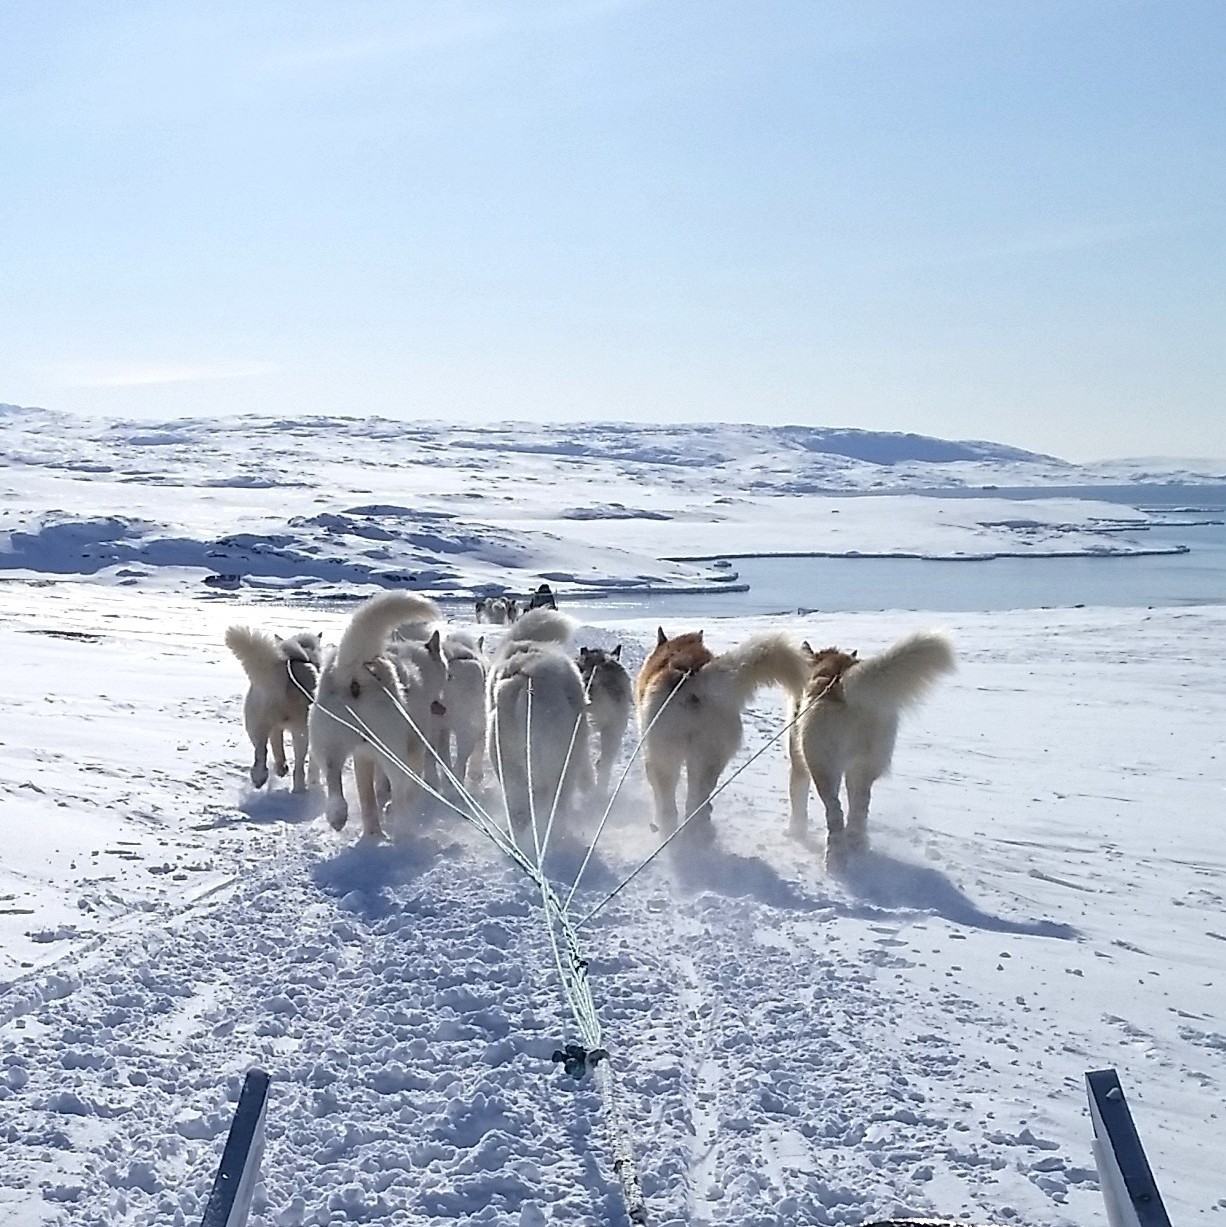
<p><img src="/assets/media/Blog_January_2026_TF_Figure.jpg" alt="Sledge dogs" title="Sledge dogs" /></p>

<p><em>Greenland Sled Dogs (Qimmit) being driven by students at the Qasigiannguit (Greenland) high school in a fan-hitch formation (Photo by Tatiana Feuerborn).</em></p>
<p> </p>

<p>For over 800 years, Greenland’s iconic sled dogs, also known as Qimmit, have been an inseparable companion to the Inuit people, pulling sleds across frozen landscapes and enabling survival in one of Earth’s harshest environments. But today, these remarkable dogs face an uncertain future. Climate change is melting the sea ice they depend on, snowmobiles are replacing dog teams, and their numbers have plummeted from 25,000 in 2002 to just around 13,000 in 2020 [<strong>1</strong>].
 </p>

<p>What makes the Qimmit unique among Arctic dog breeds? For nearly a millennium, they have served as sled dogs in the same region, partnering with the same communities. In contrast, other indigenous breeds, such as Siberian Huskies and Alaskan Malamutes, have been extensively crossbred with European dogs or adapted primarily as companions. The Qimmit, however, have remained closely tied to their original working role across Kalaallit Nunaat (Greenland). Our AaRC speaker Dr. Tatiana Feuerborn described how she joined forces with other peers to analyze Qimmit genomic data spanning from 800 years before present to present days [<strong>2</strong>]. Their findings reveal a population shaped by centuries of isolation, regional differentiation, and remarkable resilience.
 </p>

<h2 id="a-tale-of-two-arrivals">A tale of two arrivals</h2>
<p>They started by analyzing data from four different regions in Greenland: North (Avanersuaq), Northeast, East (Tunu), and West (Kitaa). Using demographic modeling, they determined that Qimmit probably arrived in Greenland in two distinct waves with the Inuit from Canada. The first wave, arriving around 1,164 years ago, gave rise to the Avanersuaq population, and the now-extinct Northeast population. A second wave, approximately 930 years ago, established the ancestral population that would split into the Kitaa and Tunu populations around 792 years ago. These findings align with archaeological evidence of a three-phase settlement of Greenland by the Inuit [<strong>3</strong>], and put Qimmit dogs as the oldest known dog breeds [<strong>4</strong>]. However, they also suggest something intriguing: either the dogs diverged before the settlement, or the Inuit arrival to Greenland occurred more than a century earlier than previously thought.
 </p>

<h2 id="the-lost-population">The lost population</h2>
<p>Nevertheless, the most poignant discovery concerns the extinct Qimmit population of Northeast Greenland. The Inuit of this region disappeared following their only recorded contact with Europeans in 1823, leaving no oral histories or human genetic legacy. The dog genomes, however, preserve their story. Dr. Feuerborn and collaborators revealed that the Northeast Qimmit were genetically highly homogeneous, evidencing a small, isolated population. When the region was resettled in 1925 by Inuit from the East, the genetic discontinuity was complete. Today’s Northeast dogs descend from these later arrivals related to the Tunu population, and not the original lineage.
 </p>

<h2 id="surviving-on-the-edge">Surviving on the edge</h2>
<p>After centuries of isolation, the Qimmit show concerning genetic trends in some regions. As Dr. Feuerborn described, the genetic diversity in Avanersuaq (North region) decreased between 1977 and the present, likely reflecting disease outbreaks including a devastating distemper epidemic that killed ~80% of dogs [<strong>2</strong>]. In Tunu (East), a similar effect was found through the 19th century, mirroring human population crashes from famine. Fortunately, 20th-century urbanization reversed this, as dogs from small isolated groups migrated to larger settlements, increasing the chances of unrelated mating. Overall, Qimmit’s genetic diversity was comparable to other wild isolated populations like dingoes, but they showed shorter homozygosity segments, indicative of a sustained small population over many generations.</p>

<h2 id="a-relatively-unaltered-lineage">A relatively unaltered lineage</h2>
<p>A surprising finding challenged assumptions about colonial influence. Despite over 300 years of Danish-Norwegian presence in Greenland, Dr. Feuerborn highlighted the minimal European dog ancestry present in modern Qimmit. This contrasts sharply with other Arctic breeds. They identified a few heavily admixed individuals, evidencing a distinct dark-furred lineage mostly used for coat fur. But these remained exceptions. The low European dog influence reflects successful early conservation policies that created a protected “sledding district” where only Qimmit dogs were allowed.</p>

<p>More intriguingly, Inuit oral traditions speak of deliberately breeding female dogs with wolves to strengthen their teams. The genetics, however, tell a more complex story. While Arctic dogs do show greater allele sharing with wolves compared to European or African breeds [<strong>5</strong>], this reflects ancient introgression that occurred before the lineage spread into North America. Additionally, wolves in Greenland are now restricted to northern regions, limiting contact with most Qimmit populations.</p>

<h2 id="a-complex-crossroads-ahead">A complex crossroads ahead</h2>
<p>The Qimmit now stand at a critical juncture. Effective population size estimates show accelerated decline over the past 150 years, and climate change threatens the very foundation of their existence, the sea ice. Shaped by intense natural and human selection for survival in the Arctic and performance as sled dogs, the Qimmit deserve our efforts to ensure they continue their partnership with the Inuit people. As Dr. Feuerborn notes in her paper, this study demonstrates <em>“the relevance of paleogenomic insight into current conversations and decisions centered around conservation and preservation of culturally significant species.”</em></p>

<p> </p>

<h2 id="references">References</h2>
<ol>
  <li><a href="https://www.science.org/doi/10.1126/science.aat9578">Sonne, C. et al. Greenland sled dogs at risk of extinction. Science 360: 1080 (2018).</a></li>
  <li><a href="https://www.science.org/doi/10.1126/science.adu1990">Feuerborn, T.R. et al. Origins and diversity of Greenland’s Qimmit revealed with genomes of ancient and modern sled dogs. Science 389: 163–168 (2025).</a></li>
  <li><a href="https://aa.uwpress.org/content/59/1/3">Mønsted, A. et al. An early Inuit workshop at a Qassi, a men’s house, Nuulliit, Northwest Greenland. Arctic Anthropol 59: 3–38 (2023).</a></li>
  <li><a href="https://www.guinnessworldrecords.com/world-records/oldest-dog-breed">Guinness World Records, “Oldest dog breed” (2020); https://www.guinnessworldrecords.com/world-records/oldest-dog-breed.html.</a></li>
  <li><a href="https://onlinelibrary.wiley.com/doi/full/10.1111/eva.12595">Pilot, M. et al. Widespread, long-term admixture between grey wolves and domestic dogs across Eurasia and its implications for the conservation status of hybrids. Evol Appl 11: 662–680 (2018).</a></li>
</ol>

<p> </p>

<p>Below, Tatiana shared with us further details about her profile, career, prospects and future projects:
 </p>

<p><strong>1. Briefly introduce yourself. What is your origin story for how you got into science?</strong> <br />
I came to ancient DNA research through a lifelong fascination with dogs. As a child, I wanted to be a veterinarian to work with animals. In school I discovered a love for history and travel. Little did I know as a child that it would be possible one day to studying archaeology and combine these interests. Today, my research focuses on the ancient DNA of Arctic dogs, where my passion for animals, curiosity about human history, and my love of travel and snow are able to come together in my work.</p>

<p><strong>2. How and/or why did you start working on this project?</strong> <br />
This project started as part of my PhD thesis that was borne out of a large project on studying the cultural and genetic origin of the Greenland Sled Dog, called the Qimmeq Project.</p>

<p><strong>3. Were there any major challenges in this project? How did you overcome them?</strong> <br />
As with all ancient DNA projects there were countless hurdles to overcome in the lab working with the samples. For me personally the biggest challenge was to do justice to the project and dogs that I cared so much about when it came to integrating a large dataset of ancient and modern genomes together and bringing results back to the communities that have cared for these dogs for millennia.</p>

<p><strong>4. What do you think are the main take-home messages of this project?</strong> <br />
I hope that this project has brought awareness of the Greenland Sled Dog to the attention of more people as these are such majestic and culturally significant dogs and shown that studying ancient populations can have tangible relevance to populations today.</p>

<p><strong>5. What do you think is missing in the field that you would like to work on?</strong> <br />
I would love to expand the research to look across the region to see in other isolated locations the unique histories of other dog populations to explore the dynamic histories of humans and their canine companions.</p>

<p><strong>6. Where do you see yourself in the near future?</strong> <br />
In the near future I see myself back in Europe expanding my research into dog history. In fact, I’m starting a new position at the University of Copenhagen and University of Greenland exploring the dogs associated with the Norse in South Greenland and the local Inuit dog populations.</p>]]></content><author><name>AaRC</name></author><category term="Blog" /><summary type="html"><![CDATA[January 2026 A genetic journey through the ancient lineage of Greenland’s Qimmit]]></summary></entry></feed>